FDA Medical Device Registration and Device Listing (US Establishment Registration)
FDA medical device registration is the yearly requirement for establishments that manufacture, sterilise, repackage, relabel, design or import medical devices and IVDs for the United States to register with the Food and Drug Administration and to list each device, under 21 CFR Part 807. Foreign establishments must also name a US Agent and pay the same annual fee as US establishments. Registration and listing are not FDA approval or clearance. This guide explains who must register, how listing works, the renewal window and what foreign manufacturers need to prepare.
Definition
What Is FDA Medical Device Registration and Listing?
FDA medical device registration has two parts. Establishment registration identifies the site and its owner or operator. Device listing identifies every device that the site makes or processes for the US market, using FDA product codes and, where one is required, the premarket submission number.
Both are entered in the Device Registration and Listing Module of the FDA Unified Registration and Listing System (FURLS). The legal basis is section 510 of the Federal Food, Drug, and Cosmetic Act and 21 CFR Part 807. The information is public and can be searched in the FDA Establishment Registration and Device Listing database.
Do you need an email containing full details within 2 minutes?
Enter your email address and we will send the medical device registration and listing service details straight away. Privacy Policy
Scope
Who Must Register, List and Pay the FDA Fee?
The FDA decides by activity. The table summarises the main establishment types from the FDA’s own list (FDA).
| Establishment type | Register | List | Pay fee |
|---|---|---|---|
| Manufacturer, including kit assemblers | Yes | Yes | Yes |
| Contract manufacturer, including contract packagers | Yes | Yes | Yes |
| Contract steriliser | Yes | Yes | Yes |
| Specification developer | Yes | Yes | Yes |
| Relabeler or repackager | Yes | Yes | Yes |
| Remanufacturer; reprocessor of single-use devices | Yes | Yes | Yes |
| Foreign exporter | Yes | Yes | Yes |
| Initial importer in the USA | Yes | No, but must identify the manufacturers | Yes |
| Manufacturer of accessories or components labelled for end-user health use | Yes | Yes | Yes |
| Manufacturer of components supplied only to finished device manufacturers | No | No | No |
| Wholesale distributor that is not a manufacturer or importer | No | No | No |
| Device used only under an investigational device exemption (IDE) | No | No | No |
In vitro diagnostic devices follow the same rules. If your product is not a device, see US FDA Registration Services for the other routes.
Outside the USA
How Do Foreign Device Manufacturers Register with the FDA?
A foreign establishment must register, list its devices and name a US Agent before its devices are exported to the USA. Under 21 CFR 807.40 the US Agent must live or keep a place of business in the USA. The agent helps the FDA communicate with the establishment, answers questions about the imported devices and helps schedule inspections.
The foreign establishment also identifies its US importers in the registration. I3CGLOBAL provides US FDA Agent Services for device and IVD establishments.
Indian medical device and IVD manufacturers
Each manufacturing site registers with its own DUNS number. Class I exempt devices can be listed straight away; devices that need a 510(k) are listed after clearance.
Manufacturers elsewhere in Asia
Contract manufacturers and contract sterilisers register in their own right, even when a US or European company owns the design and holds the 510(k).
Manufacturers in Europe
CE marking under the MDR or IVDR and an ISO 13485 certificate are not accepted in place of registration, although the quality system work supports FDA requirements.
Process
How Does FDA Medical Device Registration Work?
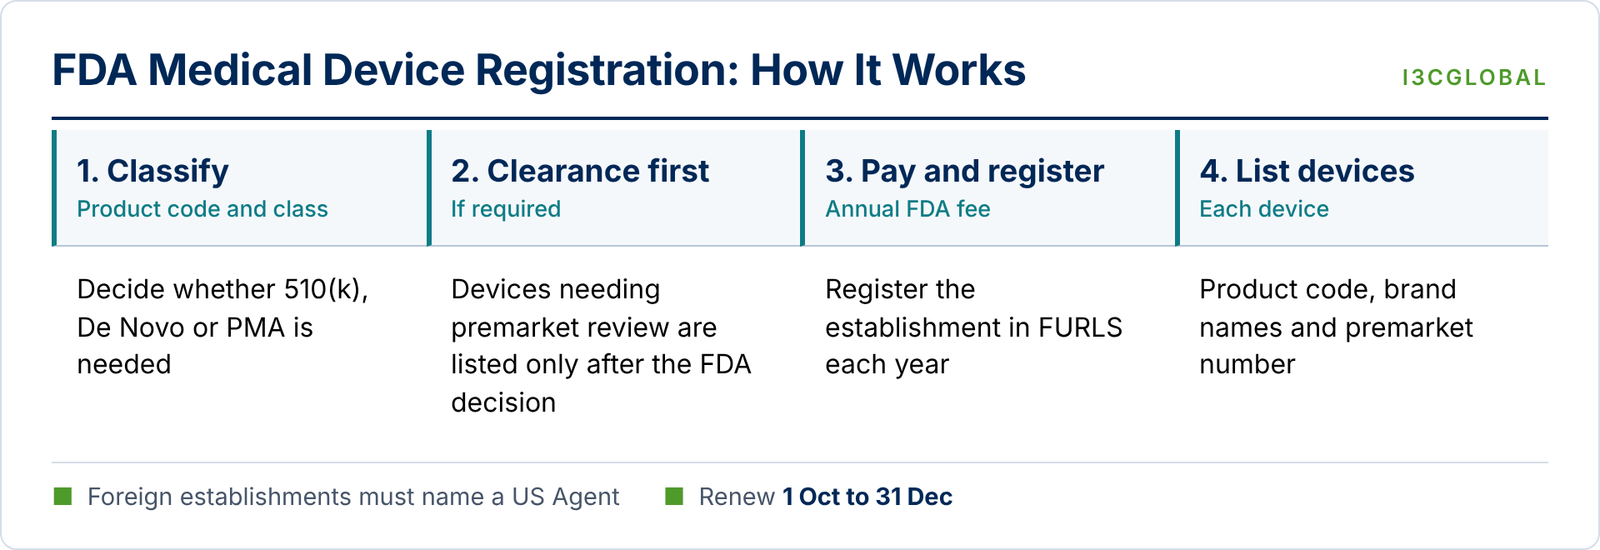
- Classify the deviceFind the FDA product code, device class and whether a 510(k), De Novo or PMA is required. See FDA 510(k).
- Obtain any premarket clearance or approvalA device that needs a premarket submission cannot be listed until the FDA has cleared or approved it.
- Appoint a US AgentForeign establishments only.
- Find and check the DUNS numberThe name and address in the DUNS database must match the registration. Our guide How to obtain a DUNS number (PDF) explains the steps.
- Pay the annual establishment registration feePayment is made on the FDA user fee website, which issues the payment identification and confirmation numbers needed for registration.
- Register the establishment in FURLSOwner or operator, official correspondent, establishment activities, US Agent and importers.
- List each deviceProduct code, proprietary names and, where it applies, the premarket submission number.
Documentation
What Information Does FDA Device Registration Require?
| Information | Notes |
|---|---|
| Establishment name, address and DUNS number | Must match the DUNS record |
| Owner or operator details | The company that owns or controls the establishment |
| Official correspondent | Contact person for the FDA |
| Establishment type | Manufacturer, contract manufacturer, steriliser, specification developer and so on |
| US Agent name, address, telephone and email | Foreign establishments |
| US importers | Foreign establishments |
| Payment identification and confirmation numbers | Proof that the annual fee has been paid |
| Product code and device class for each device | From the FDA classification database. See FDA medical device classification |
| Proprietary or brand names | All names under which the device is marketed |
| Premarket submission number | 510(k), De Novo or PMA number, or exempt status |
Maintenance
When Must FDA Medical Device Registration Be Renewed?
Registration is renewed every fiscal year between 1 October and 31 December (FDA). The fee for the new fiscal year is paid first. During renewal the establishment reviews every listing and confirms that it is still correct.
| Event | What to do | When |
|---|---|---|
| Annual renewal | Pay the fee, then certify the registration and listings | 1 October to 31 December |
| New device | Add a listing | Before the device is commercially distributed |
| Device discontinued | Remove or deactivate the listing | At the next update |
| Change of US Agent, importer or official correspondent | Update the registration | When the change takes effect |
| Change of ownership or site address | Update or re-register as required | When the change takes effect |
Current FDA and service fees are on our FDA medical device registration fee page.
Important distinction
Is FDA Device Registration the Same as Clearance or Approval?
No. The FDA states that a registration and listing entry “does not denote approval, clearance or authorization” of a facility or its devices, and that it “does not issue registration certificates to medical device facilities” (FDA).
An owner or operator number, a registration number and public database entries for the establishment and its devices.
510(k) clearance, De Novo classification, PMA approval, an FDA certificate or the right to use the FDA logo. See FDA Registration Certificate.
Registered establishments must also meet the Quality Management System Regulation in 21 CFR Part 820 and the labeling rules in 21 CFR Part 801. See FDA QMSR, FDA Labeling Requirements and the FDA Approval Process.
Most devices sold in the US also need a Unique Device Identifier (UDI) on the label and an entry in the FDA GUDID database. This is a separate requirement from registration and listing. See UDI system.
Avoidable problems
What Are the Common Device Registration Mistakes?
- Listing a device before clearance. A device that needs a 510(k) cannot be listed until the FDA has cleared it.
- Advertising “FDA registered” as if it meant approved. The FDA treats this as misleading.
- Wrong establishment type. A design owner that outsources production is a specification developer, not a manufacturer.
- Forgetting the contract steriliser or contract manufacturer. Each site registers separately.
- Paying the fee late. The registration cannot be renewed without the payment confirmation number.
- Using the US distributor as US Agent. Changing distributor then leaves the registration without an agent.
Work with I3CGLOBAL
How Does I3CGLOBAL Handle FDA Medical Device Registration?
- Device classification and product code check by our US FDA consultants
- US Agent service for device and IVD establishments
- DUNS record check and FURLS account set-up
- Establishment registration and device listing
- Annual renewal reminders and listing updates
Start your medical device registration
Send us the device description, intended use and site details.
FAQs
FDA Medical Device Registration: Frequently Asked Questions
What is FDA medical device registration?
It is the annual registration with the US Food and Drug Administration of establishments that manufacture, sterilise, repackage, relabel, develop specifications for or import medical devices for the United States, together with a listing of the devices they handle. It is required by section 510 of the FD&C Act and 21 CFR Part 807.
Is FDA establishment registration the same as device listing?
No. Establishment registration identifies the site and its owner or operator. Device listing identifies each device made or processed there by product code, and for devices that need it, by the 510(k), De Novo or PMA number.
Do foreign medical device manufacturers need to register with the FDA?
Yes. Foreign manufacturers, contract manufacturers, contract sterilisers, specification developers and foreign exporters must register, list their devices, pay the annual fee and name a US Agent.
When must FDA medical device registration be renewed?
Every year between 1 October and 31 December, for the fiscal year that started on 1 October. The annual establishment registration fee is paid before the renewal is submitted.
Does FDA registration mean a medical device is FDA approved or cleared?
No. The FDA states that registration and listing do not denote approval, clearance or authorization of a facility or its devices. Devices that need a 510(k), De Novo or PMA must obtain it separately before they are listed and sold.
Does the FDA issue a medical device registration certificate?
No. The FDA does not issue registration certificates to medical device facilities. Registration can be checked in the FDA’s public Establishment Registration and Device Listing database.
Do IVD manufacturers register in the same way?
Yes. In vitro diagnostic devices are medical devices under US law, so IVD manufacturers register the establishment and list each IVD under its product code.
Do component suppliers have to register?
A manufacturer of components that are sold only to finished device manufacturers does not register. A manufacturer of accessories or components that are packaged and labelled for health use by the end user does.