
IVDR Consultants
I3CGlobal’s IVDR consultants help in vitro diagnostic manufacturers move from classification to notified body certificate under Regulation (EU) 2017/746. We cover every risk class, from self-declared Class A to Class D, and prepare the technical documentation, performance evaluation and QMS evidence that notified bodies expect. IVDR is one of our core specialties, with regulatory and product technical support delivered as one service.
Why IVD Manufacturers Work with IVDR Consultants
The EU IVDR (Regulation (EU) 2017/746) has applied since 26 May 2022 and changed the rules far more than the old IVD Directive 98/79/EC. Around 80% of IVDs now need a notified body, against roughly 20% before, and every device needs a performance evaluation, a post-market surveillance system and IVDR-format technical documentation.
Most manufacturers have the product knowledge but not the time or the regulatory experience to rebuild their files. IVDR consultants close that gap. We have helped manufacturers certify under the IVDD and now support them through the IVDR transition, so we know where legacy files fall short: clinical data, the performance evaluation report (PER) and post-market surveillance.
Is your current technical documentation ready for a notified body? If you are not sure, a gap assessment is the fastest way to find out.
IVDR Consulting Services
Our IVDR consulting covers the full project, or any single deliverable your team needs help with. Typical scope includes:
Due diligence & gap assessment
Review of your existing IVDD or draft IVDR files against Annexes I, II and III, with a prioritised gap list and plan.
Classification & intended purpose
Confirmation of the device class under Annex VIII and review of the intended purpose wording that drives it. See IVDR classification.
Technical documentation
Guidance on and preparation of the Annex II and III technical documentation file, including design files, approval records and verification and validation reports.
GSPR & risk management
Documenting compliance with the General Safety and Performance Requirements and reviewing your ISO 14971 risk management file.
Performance evaluation
Performance evaluation plan, scientific validity, analytical performance and clinical performance, compiled in a PER. We work with associated laboratories for performance studies.
Post-market surveillance
PMS plan, PMS report or PSUR, and the post-market performance follow-up plan and report.
QMS upgrade
Upgrading your EN ISO 13485 system to IVDR requirements, including PRRC, UDI and vigilance procedures. See QMS upgrade to IVDR.
EUDAMED & EU representative
Actor and device registration in EUDAMED, UDI assignment and EU authorised representative support for non-EU manufacturers.
Notified body support
Notified body selection, application, technical review responses and file updates until the certificate is issued.
IVDR Consultants for Class A, B, C and D Devices
The work and the evidence grow with the risk class. We scope each project to the class and route that apply.
| Class | Notified body | Where we focus |
|---|---|---|
| Class A | No: self-declaration | Technical documentation, performance evaluation, labelling and EUDAMED |
| Class A sterile | Sterility aspects only | Sterilisation, packaging and shelf-life evidence, plus the full Class A file |
| Class B | Yes: sampled per device category | Device grouping, audit-ready files for every device, QMS upgrade |
| Class C | Yes: per generic device group | Clinical performance evidence, PER, SSP and PSUR |
| Class D | Yes: every device, plus EURL and expert panel | Common specifications, batch verification and expert panel readiness |
How Our IVDR Consultants Run a Project
- Gap assessmentWe review your current documentation, QMS and evidence and agree the scope, class and timeline.
- Plan the evidenceWe define the performance evaluation strategy, studies needed and the technical documentation structure.
- Build the documentationOur consultants prepare the technical file, GSPR checklist, PER and PMS documents with your technical team.
- Upgrade the QMSWe align your ISO 13485 system with the IVDR and prepare you for the notified body audit.
- Notified body submissionWe support the application, answer technical review questions and update files until certification.
- After the certificateWe help maintain PMS, PMPF, PSUR and technical documentation updates through surveillance.
Why Choose I3CGlobal as Your IVDR Consultants
All risk classes
We take up IVD projects of every class and device type, from instruments and specimen receptacles to Class D blood screening assays.
Regulatory + technical in one team
Regulatory documentation and product technical consultation are delivered together, so evidence and documents match.
Certified ways of working
Our QMS is certified to ISO 13485 and our data security to ISO 27001, so your technical data is handled under controlled processes.
Fast responses
We reply to technical emails within 24 hours, which keeps notified body review rounds short.
Laboratory network
We work with associated laboratories for analytical and clinical performance studies.
Global support
Our team supports IVDR projects for manufacturers in every region, with an EU authorised representative where needed.
IVDR Consulting Cost
IVDR consulting cost varies widely. It depends mainly on:
- The project scope: full technical documentation or selected deliverables
- The risk class and novelty of the IVD
- The complexity of the intended use
- The number of device models, variants and device groups
- How much performance data already exists
A typical IVDR technical documentation project takes around 3 to 4 months, depending on how quickly product and scientific information reaches our consultants. Notified body timelines come on top of that.
See indicative budgets on our IVDR CE certification cost page, or estimate your own with the IVDR cost calculator.
Resources for your team
- IVDR technical documentation support
- Risk analysis procedure & templates, useful for small and medium-sized manufacturers
- IVDR transition timeline
- IVD CE marking process
Do you need an email containing full details about our IVDR consulting services within 2 minutes? Share your email below. Privacy Policy
Have an IVDR Transition Strategy in Place?
If not, don’t worry. Our IVDR consultants and regulatory experts have the qualifications and experience to take on IVDR projects of any risk class, from anywhere in the world.
- Gap assessment and IVDR transition plan
- Technical documentation and performance evaluation
- QMS upgrade, PRRC and EUDAMED support
- Notified body application and review support
Start your IVDR project
Share your device details and get a tailored proposal.
IVDR Consultants: Frequently Asked Questions
What do IVDR consultants do?
IVDR consultants help manufacturers meet Regulation (EU) 2017/746: confirming the class, preparing technical documentation and the performance evaluation, upgrading the QMS, registering in EUDAMED and supporting the notified body review until certification.
Do you provide assurance of CE marking?
We offer several service types with matching pricing options. With our assured service, we guarantee CE marking for your project.
How long does IVDR technical documentation take?
Typically around 3 to 4 months, depending on how promptly product technical and scientific information is delivered to our IVDR consultants. Notified body review time is additional.
Can you help with performance studies?
Yes. We plan analytical and clinical performance studies and work with associated laboratories to run them, then compile the results in the performance evaluation report.
Do you support manufacturers outside the EU?
Yes. We support IVD manufacturers worldwide and can provide EU authorised representative services and EUDAMED registration.
Can I3CGlobal support both regulatory documentation and product technical consultation?
Yes. We provide IVDR regulatory and product technical consultation as a combined service, from classification to notified body certificate.
What are the IVDR deadlines for legacy devices?
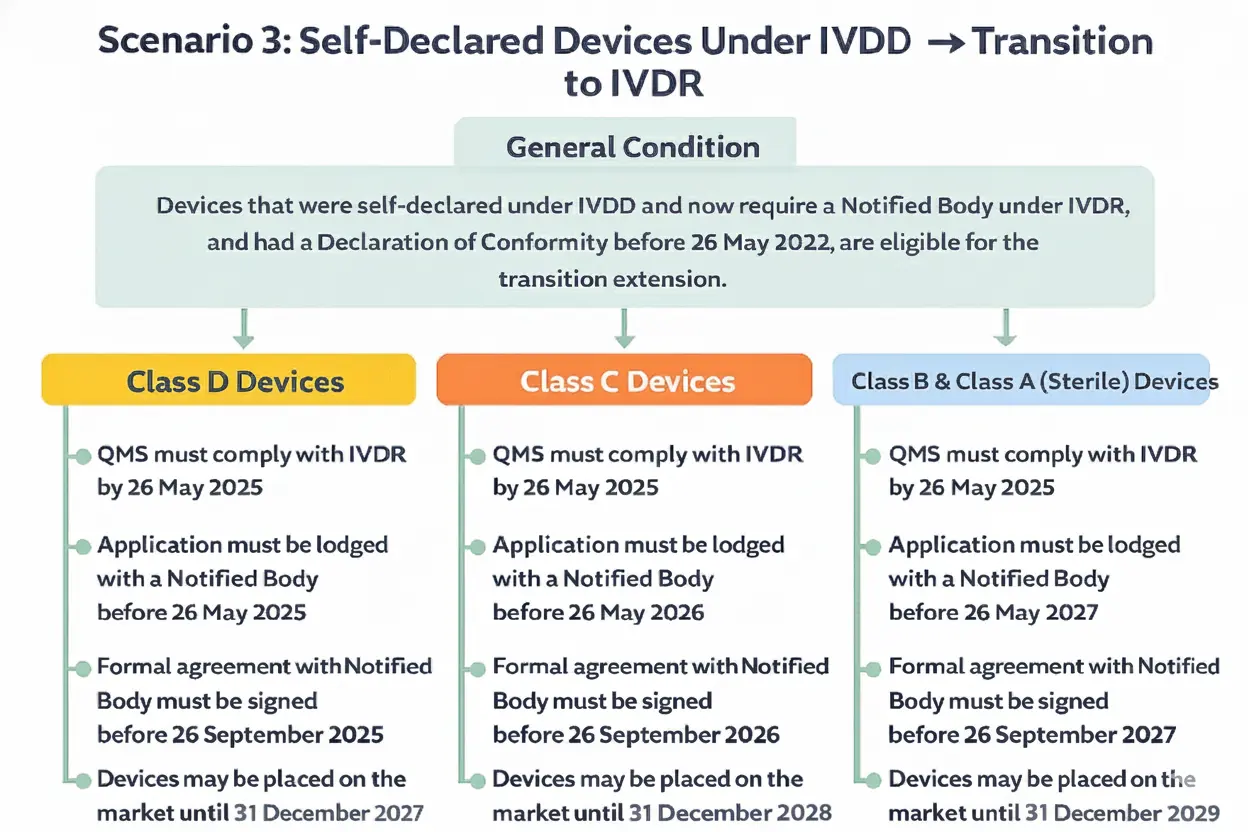
Eligible legacy devices have until 31 December 2027 (Class D), 31 December 2028 (Class C) or 31 December 2029 (Class B and Class A sterile), provided the conditions of Regulation (EU) 2024/1860 are met.
Does a consultant replace the notified body or authorised representative?
No. A notified body independently assesses conformity. An EU authorised representative is a legal role for non-EU manufacturers. A consultant helps you prepare and manage the work.
Do Class A devices need a notified body?
Non-sterile Class A devices are self-declared. For Class A sterile devices, the notified body’s assessment is limited to the sterile aspects.
What do you need to provide for a quote?
Share the intended purpose, the expected class, the number of device variants, your current QMS status and the performance data you already have.
Updated | Author: Sobha Augustine, Founder, I3CGLOBAL | Reviewed by: Swathi Tummala, Senior Regulatory Consultant, I3CGLOBAL