
MDR Article 117 and the Notified Body Opinion
MDR Article 117 amends Annex I of Directive 2001/83/EC. If an integral drug-device combination has no EU declaration of conformity or notified body certificate for its device part, and that device would need a notified body if used separately, the marketing authorisation application must include a notified body opinion (NBOp). I3CGLOBAL’s MDR Article 117 consultants help applicants prepare for it.
What Is MDR Article 117?
MDR Article 117 is the provision of Regulation (EU) 2017/745 that amends section 3.2, point 12 of Annex I to Directive 2001/83/EC. It links medicines law to device law for products where the drug and the device form one integral product.
These products are authorised as medicinal products, so the European Medicines Agency (EMA) or a national competent authority remains the decision maker. However, the authority does not assess conformity of the device part. The dossier must therefore show that the device part meets the relevant General Safety and Performance Requirements (GSPRs) in Annex I of the MDR.
In practice, MDR Article 117 asks one question: is there evidence of device conformity in the dossier? If the evidence is a declaration of conformity or an EU certificate, no opinion is needed. If neither exists and a notified body would normally be involved, the applicant must obtain an NBOp.
Key facts
MDR Article 117 at a glance
- Legal basis
- Regulation (EU) 2017/745, Article 117
- Amends
- Directive 2001/83/EC, Annex I, section 3.2, point 12
- Applies from
- 26 May 2021
- Products
- Integral products under MDR Article 1(8) and 1(9), second subparagraphs
- Assessment
- Device part against the relevant GSPRs in MDR Annex I
- Output
- Notified body opinion (NBOp), submitted with the marketing authorisation dossier
- Decision maker
- EMA/CHMP or the national competent authority
Which Products Fall Under MDR Article 117?
MDR Article 117 covers integral drug-device combinations. The MDR describes two routes into this group, and in both the product is governed as a medicinal product.
1(8)
Device incorporating a medicinal substance
The device incorporates, as an integral part, a substance that would be a medicinal product if used separately, and the action of that substance is principal. EMA’s example is an ingestible sensor.
1(9)
Device administering a medicinal product
The device and the medicinal product form a single integral product, intended only for use in that combination, and it is not reusable. Examples include pre-filled syringes, pens, inhalers, sprays and transdermal patches.
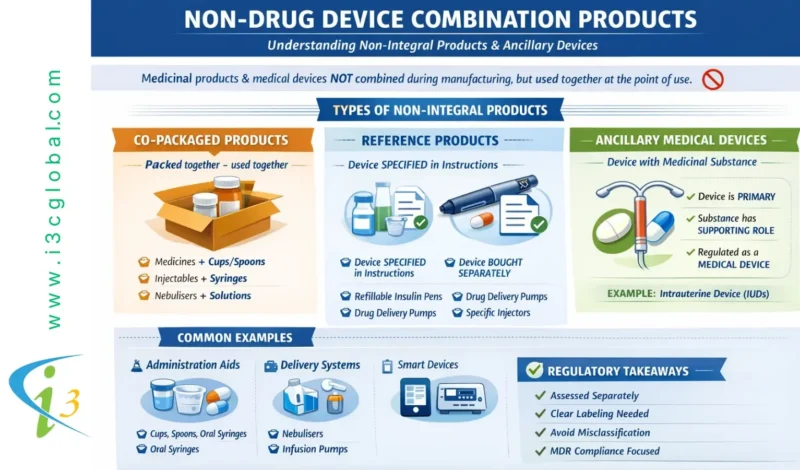
How do integral products differ from other combinations?
Classification decides the regulatory route, so confirm it before planning any notified body work. The table compares the main product types.
| Product type | What it means | Examples | Route |
|---|---|---|---|
| Integral | Drug and device form a single product with a principal medicinal action or single-use combined administration | Pre-filled syringes, single-use pre-filled inhalers | Medicinal product authorisation, with MDR Article 117 evidence |
| Co-packaged | Medicinal product and device are placed on the market together in one pack, but are not integrated | Oral syringe or dosing cup in the pack | Not an MDR Article 117 product; EMA covers these separately |
| Referenced | Product information refers to a specific device that is obtained separately | Refillable pens, nebulisers, pumps | Device regulated separately under the MDR |
| Ancillary substance | Device incorporates a medicinal substance, but the action of the substance is ancillary to the device | Drug-eluting stents, heparin-coated catheters | Device CE marking, with medicines authority consultation under MDR Annex IX, section 5.2 |
Excluded: combined advanced therapy medicinal products are outside MDR Article 117. Container closures without an administration function are assessed under other rules. Refer to EMA’s Q&A, Rev. 6, questions 2.1 and 2.2.
When Is a Notified Body Opinion Required?
An NBOp is required when the dossier contains no conformity result for the device part and that device, if used separately, would need notified body involvement. EMA sets out the expected evidence by device class in Table 1 of its Q&A.
| Device part, if used separately | Expected evidence | If the evidence is not available |
|---|---|---|
| Class I, not sterile, without measuring function and not a reusable surgical instrument | EU declaration of conformity | Statement of compliance with the relevant GSPRs from the marketing authorisation holder |
| Class I sterile, measuring or reusable surgical instrument | EU certificate from a notified body designated for that device type | NBOp required |
| Class IIa, IIb or III | EU certificate from a notified body designated for that device type | NBOp required |
What about products authorised before 26 May 2021?
MDR Article 117 is not retrospective. It does not apply to integral products already authorised, or to applications submitted, before 26 May 2021. A major device change or a new device after that date still needs the relevant declaration, certificate or opinion.
- Certificates issued under Directives 90/385/EEC or 93/42/EEC may support the application while they remain valid under the MDR transitional provisions.
- EMA states that class I declarations drawn up before 26 May 2021 may be accepted until 31 December 2028.
- The evidence should ideally be filed with the application. Late documents must arrive before the opinion on the medicinal product, and may cause additional clock stops.
What Documents Does the Notified Body Assess?
The notified body reviews technical documentation for the device part. Team-NB’s MDR Article 117 position paper groups the file into three parts.
A
Device description
Intended purpose, users, patient population, indications, contraindications, principles of operation, key functional elements, materials and technical specifications.
B
Instructions for use
The instructions for use or package insert for the integral product.
C
GSPR documentation
The GSPR list, justification for those that do not apply, conformity methods and results, detailed evidence, standards applied and the controlled documents that hold the evidence.
Applicants usually draw this evidence from the device’s technical file, ISO 14971 risk management, usability engineering, sterilisation validation and clinical evaluation, where each applies.
What does the notified body opinion cover?
The opinion addresses the conformity of the device part with the relevant GSPRs. It is not a CE certificate. Team-NB’s opinion template records the product, the review type, the GSPR assessment by chapter, the solutions adopted, recommendations to the competent authority, standards and the data reviewed.
The notified body may also consider how the medicinal product affects device safety and performance, for example on compatibility or sterilisation. The EMA or national authority assesses the medicinal product itself, using its quality guideline for products used with a device.
Partial opinions are not accepted: EMA expects the opinion to confirm full compliance with the relevant GSPRs, and its scope must match the intended purpose in the application.
How Do You Obtain an NBOp Under MDR Article 117?
These seven steps give a practical sequence. Exact requirements vary by notified body, so agree the details early with the one you select.
- Confirm the product typeEstablish whether the product is integral, co-packaged, referenced or ancillary, and document the principal mode of action.
- Classify the device partApply the MDR Annex VIII rules as if the device were used separately. This shows whether notified body involvement is needed.
- Check existing evidenceLook for a valid declaration of conformity or EU certificate. If one exists, an NBOp may not be needed.
- Close the GSPR gapsComplete the GSPR checklist and generate missing evidence, such as biological evaluation, risk analysis, usability and packaging data.
- Select a notified bodyChoose a body designated under the MDR for the device type, and confirm that it accepts Article 117 applications. Check its fees and capacity.
- Submit and answer questionsProvide a legible, searchable and bookmarked file with no access protection. Track each question from the reviewer until it is closed.
- File the opinion in the dossierInclude the issued opinion in the marketing authorisation application, and keep it aligned with the device described in the dossier throughout the review.
How Long Does a Notified Body Opinion Take?
The documents reviewed for this page give no fixed review period for an NBOp. Timing depends on the notified body, the quality of the file and the number of questions raised.
As one published planning figure, PharmaLex estimates 2 to 3 months for the review, plus clock stops for questions. Treat this as an estimate, and confirm the timing and fees with your notified body before fixing a submission date.
Evidence generation often takes longer than the review. New biocompatibility, usability or stability work should therefore be scheduled well before the marketing authorisation submission.
Which Changes Need a New Notified Body Opinion?
MDR Article 117 continues to apply after authorisation. EMA expects a new or updated opinion when a new device is introduced, for major device changes, and for medicinal product changes that may affect device performance. Variations follow the current EU variations guidelines, which EMA notes apply from 15 January 2026.
Team-NB’s lifecycle position paper offers typical examples, and the list is not exhaustive.
| Typically substantial | Typically non-substantial |
|---|---|
| Change to intended purpose, operating principle or sterilisation method | Tightening specifications within the assessed range |
| New material or component that does not meet the assessed specification | Component that meets the same assessed specification |
| Software change that may affect diagnosis or therapy | Bug fixes or user interface changes with no therapeutic effect |
| Packaging change that affects function, safety, stability or seal integrity | Shelf-life change validated by an assessed protocol, except implantable or ingestible devices |
The marketing authorisation holder analyses each change and makes the final decision. Where no new opinion is obtained for a variation, the holder must justify this in Module 3.2.R. Agreements with the device manufacturer should also cover how device changes are communicated, as device post-market data can trigger them.
MDR Article 117 Support from I3CGLOBAL
I3CGLOBAL is a global regulatory consulting company for medical devices, IVDs, pharmaceuticals and life sciences. Our QARA team helps applicants prepare technical documentation, work with notified bodies and meet MDR Article 117 requirements. The notified body issues the opinion, and the medicines authority makes the authorisation decision.
| Activity | What our consultants do |
|---|---|
| 1. Identify the product | Determine whether the product is a drug-device combination or an integral product. |
| 2. Define function | Identify the main and ancillary components, the intended use and how the product works. |
| 3. Set the strategy | Use pharmaceutical and device expertise to plan the regulatory strategy and route. |
| 4. Classify the device | Check the risk class of the device part and the classification rule that applies. |
| 5. Select standards | Identify the relevant harmonised and non-harmonised standards. |
| 6. Biological evaluation | Plan and review biological evaluation as required. |
| 7. Risk analysis | Carry out risk analysis and benefit-risk assessment for the combined product. |
| 8. Clinical evaluation | Prepare and review the clinical evaluation of the device part. |
| 9. Supporting data | Review usability, cleanliness, packaging and shipping requirements. |
| 10. GSPR file | Compile and organise documents in line with the GSPRs. |
| 11. Notified body choice | Help the manufacturer find and choose a suitable notified body. |
| 12. Application | Support the application and submission to the notified body. |
| 13. Questions | Coordinate communication and answer notified body questions during the technical review. |
| 14. Document control | Track documents, make changes and add new ones during the opinion process. |
Planning an MDR Article 117 submission?
Share your product details and get a tailored proposal for your NBOp project.
MDR Article 117: Frequently Asked Questions
What is MDR Article 117 in simple terms?
MDR Article 117 requires a marketing authorisation dossier for an integral drug-device product to show that the device part meets the relevant GSPRs. Where no declaration of conformity or EU certificate is available and a notified body would be needed, the applicant must provide a notified body opinion.
What is an NBOp?
An NBOp is a notified body opinion on the conformity of the device part of an integral product with the relevant GSPRs in MDR Annex I. It supports the marketing authorisation application. It is not a CE certificate.
Which documents are required by the notified body?
The technical file should include the device description, instructions for use and GSPR documentation with supporting evidence. Team-NB describes the expected structure in its position paper on documentation requirements, which draws on MDR Annex II. Each notified body may add its own requirements.
What is the timeline for a notified body opinion letter?
The documents reviewed here give no fixed review period. PharmaLex estimates 2 to 3 months plus clock stops for questions, but this is only a planning figure. Confirm the timing with your notified body, and allow extra time to generate any missing evidence.
Does a class I device part need a notified body opinion?
Usually not, if the device is class I and not sterile, has no measuring function and is not a reusable surgical instrument. A declaration of conformity is expected, or a GSPR compliance statement from the marketing authorisation holder. Class I sterile, measuring and reusable surgical devices need notified body evidence.
Does MDR Article 117 apply to co-packaged products?
No. MDR Article 117 applies to integral products. Co-packaged medicinal products and devices are placed on the market together but are not integrated, and EMA addresses them in a separate question of its guidance.
Does MDR Article 117 apply to products already on the market?
It does not apply retrospectively to products authorised, or applications submitted, before 26 May 2021. However, it applies after authorisation when a new device is introduced or a major device change is made, so existing products may still need an opinion later.
Who is responsible for obtaining the opinion?
The marketing authorisation applicant or holder is responsible for including the evidence in the dossier. In practice, the applicant often works with the device manufacturer to provide the technical documentation, and their agreement should set out how each party handles device information and changes.
What happens if there is no notified body opinion?
Where an opinion is required, the authority will ask for it. EMA states that missing documents should be provided before an opinion on the medicinal product is issued, and their absence may cause additional clock stops.
Is the notified body opinion published?
EMA states that the opinion itself is not published separately. The European Public Assessment Report summarises the device information, including whether a declaration, certificate, opinion or statement was submitted.
Official References and Related Guides
- Regulation (EU) 2017/745 (MDR), Articles 1(8), 1(9) and 117
- Directive 2001/83/EC, Annex I, section 3.2, point 12
- EMA Q&A on implementing the MDR and IVDR, Rev. 6 (December 2025)
- EMA guideline on quality documentation for medicinal products used with a medical device
- EMA presentation: notified body opinion, the assessor’s view
- Team-NB: documentation requirements, substantial changes and opinion template
- Medical Device Technical File
- GSPR Checklist
- Clinical Evaluation Report
- Post-Market Surveillance
- MDR Article 17
- CE Certification Cost
Updated 4 November 2026 | Author: Sobha Augustine, Founder, I3CGLOBAL | Reviewed by: Asha Johnson, Manager, I3CGLOBAL