
IVDR Technical Documentation Services
IVDR technical documentation is the evidence file that shows your in vitro diagnostic device meets Regulation (EU) 2017/746. It must follow Annex II and Annex III, be available to authorities and the notified body on request, and be kept up to date for the full device lifecycle. I3CGlobal handles IVDR technical documentation preparation, review and remediation for Class A to Class D devices.
What Is IVDR Technical Documentation?
IVDR technical documentation contains all the supporting information about your IVD medical device: what it is, how it is designed and made, how it performs and how its risks are controlled. Every manufacturer placing an in vitro diagnostic device on the EU market must prepare technical documentation for IVDR compliance, regardless of risk class or notified body (IVDR Article 10(4)). Our IVDR technical documentation checklist lists every section the notified body expects.
Depending on the device's classification and the conformity assessment route chosen, a notified body may review one or more technical documentation files. The content and format are defined in Annex II and Annex III of the IVDR, Regulation (EU) 2017/746, and are based on the STED format developed by the Global Harmonization Task Force (GHTF).
A current version must be available to competent authorities on request, so it should also be held by your EU Authorised Representative (Article 11(3)) and overseen by the person responsible for regulatory compliance (Article 15). After certification, the file must be kept up to date at all times (Annex IX, Chapter II).
IVDR Technical Documentation Requirements
These are the IVDR technical documentation requirements every in vitro diagnostic device must meet, whatever its class, with each element under its own heading. IVDR Annex II and Annex III documentation is set out below: Annex II covers the device file in six chapters and Annex III adds the post-market surveillance documentation. For a section-by-section table of contents, see our IVDR technical documentation checklist.
| Section | What to include | Reference |
|---|---|---|
| Device description and specification | Intended purpose, variants, accessories, reference to previous and similar generations | Annex II, 1 |
| Information supplied by the manufacturer | Labels, IFU and any promotional material that makes claims | Annex II, 2 |
| Design and manufacturing information | Design stages, manufacturing processes, sites and critical suppliers | Annex II, 3 |
| General safety and performance requirements | GSPR checklist, applied standards and common specifications, evidence for each requirement | Annex II, 4 |
| Benefit-risk analysis and risk management | Risk management file showing risks are identified, controlled and acceptable | Annex II, 5 |
| Product verification and validation | Analytical performance, clinical performance, scientific validity, stability, usability and device lifetime, plus sterilisation, packaging and software where relevant | Annex II, 6 |
| Post-market surveillance plan | Plan to collect and assess market data, including the PMPF plan | Annex III, 1 |
| PMSR or PSUR | PMS report for Class A and B; periodic safety update report for Class C and D | Annex III, 2 |
| Vigilance | Serious incident reporting and field safety corrective actions | Articles 82 to 87 |
IVDR Technical Documentation and GSPR
The technical documentation and the performance evaluation report together must prove compliance with the General Safety and Performance Requirements in Annex I. Notified bodies look closely at these elements.
GSPR checklist and standards
Show how each requirement is met, with the harmonised standards and common specifications you applied identified clearly.
Risk management
Annex I, Section 3 treats risk management as a continuous process across the whole lifecycle that needs regular, systematic updating.
Post-market surveillance plan
Annex III, Section 1(a) requires a plan to collect and assess market data, covering serious incidents, complaints and information on similar devices.
PMPF plan
The post-market performance follow-up plan is part of the PMS plan and is driven by the results of risk management and performance evaluation.
PMSR or PSUR
Class A and B devices need a PMS report. Class C and D devices need a PSUR for each device, updated at least annually and submitted to the notified body.
Performance evaluation report
The PER under Annex XIII brings together scientific validity, analytical and clinical performance, and must be submitted with the technical documentation.
IVDR Technical Documentation Preparation: How We Work
- Classification and routeConfirm the IVDR class and conformity assessment route for each device and variant.
- Technical documentation reviewAn IVDR technical documentation review of your existing IVDD file or design records against Annex II and Annex III, with a prioritised gap list.
- GSPR and standards mappingBuild the GSPR checklist and link each requirement to standards and evidence.
- Performance evidencePlan and document scientific validity, analytical and clinical performance, and stability.
- Compile the fileWrite the technical documentation in STED format, with risk management, labelling and PMS documents.
- Notified body supportSupport submission and answer notified body questions until the certificate is issued.
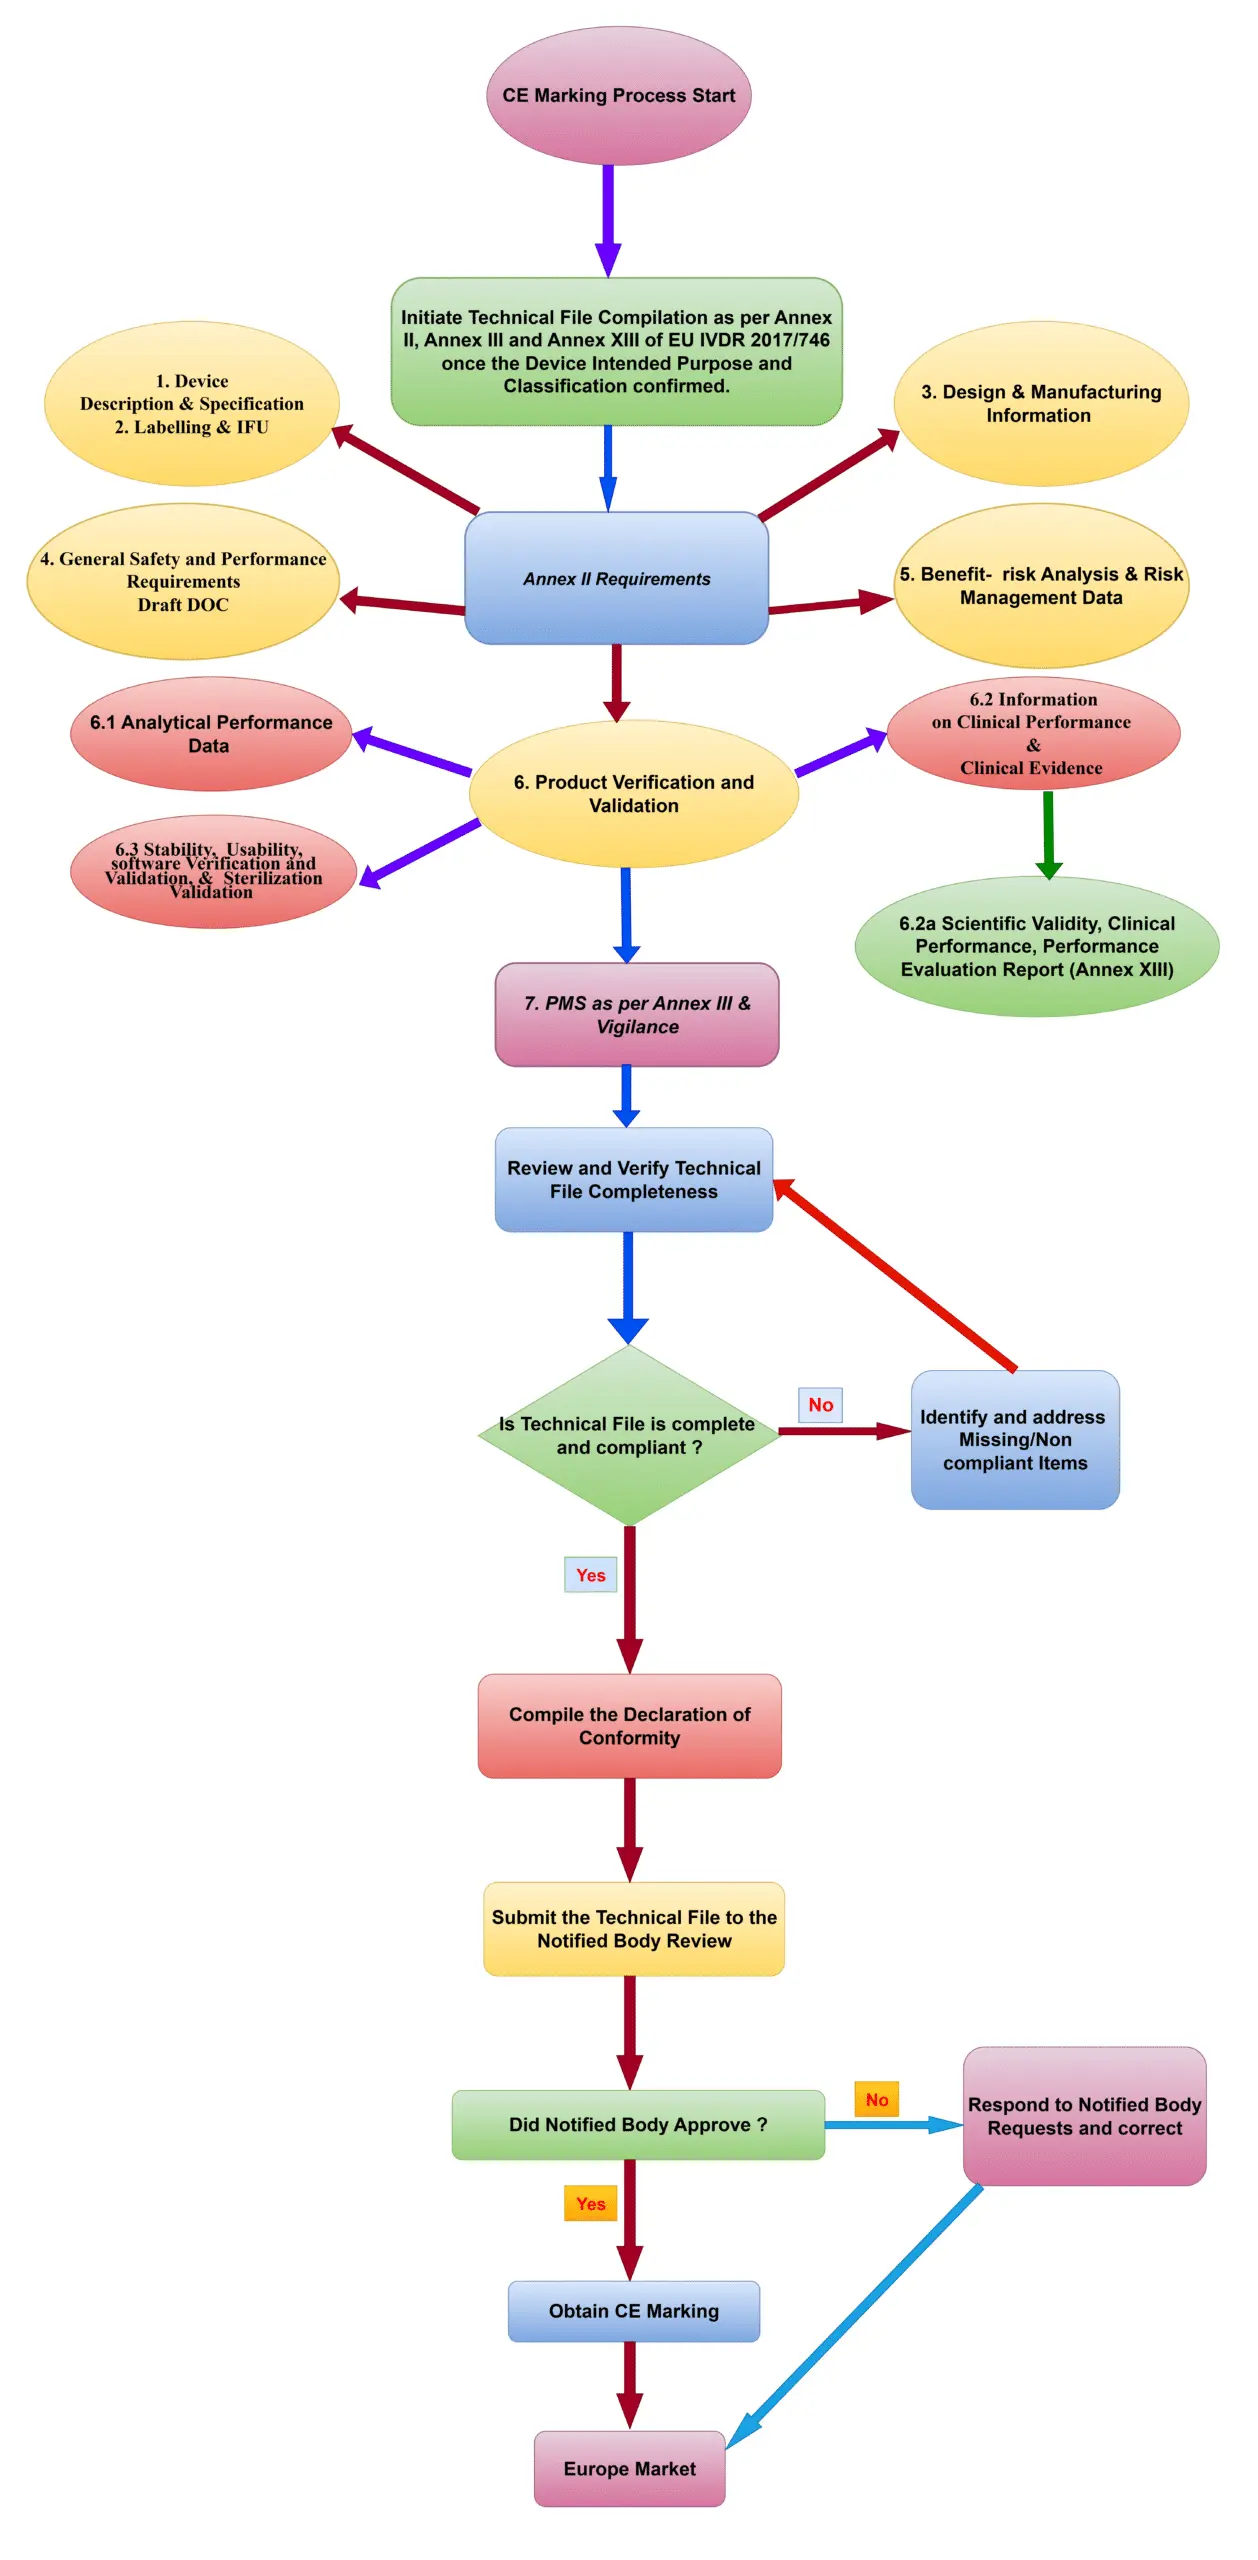
We prepare IVDR technical documentation in the Annex II and Annex III structure, so the notified body can find every piece of evidence quickly. The flowchart below shows how a typical technical file for a Class B IVD is compiled, checked for completeness, submitted and maintained until CE marking.
Technical Documentation for IVDR by Device Class
Every device class needs the same Annex II and Annex III content, but the way the notified body assesses the file changes with the risk class. The IVDR classification rules in Annex VIII decide which conformity assessment route applies, so confirm the class of each device before you plan your technical documentation for IVDR certification.
For Class B and Class C devices, the notified body assesses technical documentation on a sampling basis, so every file in your portfolio must be complete and consistent, not just the one selected first. For Class D devices, the notified body assesses the technical documentation of each device, and EU reference laboratories and expert panels may also be involved.
| Class | Notified body involvement | What happens to the technical documentation |
|---|---|---|
| Class A (non-sterile) | None, self-declaration | The manufacturer keeps the full file and makes it available to competent authorities on request. |
| Class A (sterile) | Limited to sterility | The notified body reviews the aspects related to establishing, securing and maintaining sterile conditions. |
| Class B | QMS audit and sampling | Technical documentation of at least one representative device per category of devices is assessed (Annex IX, 2.3). |
| Class C | QMS audit and sampling | Technical documentation of at least one representative device per generic device group is assessed. Self-tests, near-patient tests and companion diagnostics get additional review. |
| Class D | Every device | The file of each device is assessed, with EU reference laboratory verification and expert panel consultation where applicable (Article 48), plus batch verification. |
Key Points for IVD Manufacturers
Required for every class
Class A self-declared devices need full technical documentation too, even without notified body review.
Based on GHTF STED
The Annex II structure follows the Summary Technical Documentation format, which makes reviews faster when followed closely.
Keep it live
The file must be updated after certification as PMS data, risk files and standards change.
Real-time stability data
Stability of the device and of the specimen are both required; accelerated data alone does not support shelf-life claims.
Claims need evidence
Every claim on labels, IFU or marketing material must be supported in the file (Article 7).
IVDD experience is not enough
IVDR expectations go well beyond IVDD, so experienced IVDR consultants save review cycles.
Common Gaps in IVDR Technical Documentation
Most notified body questions trace back to a small number of recurring gaps. Fixing them before submission is the fastest way to shorten the IVDR technical documentation review and avoid extra review rounds.
Vague intended purpose
Annex II, 1.1 expects the analyte, function, disorder or condition, specimen type, testing population and intended user to be stated precisely. Gaps here spread into the performance evaluation and labelling.
Weak GSPR traceability
Checklists that only say "complies" are not enough. Each requirement should point to a specific report, document number and version.
Claims not matched by data
Analytical and clinical performance studies must use the same specimen types, population and cut-offs that the IFU claims.
Outdated standards
Superseded versions of standards such as EN ISO 14971 or EN ISO 13485 need a gap assessment or a justification for continued use.
Inconsistent documents
Device names, variants, UDI-DI and version numbers often differ between the labels, IFU, risk file and performance evaluation report.
Disconnected PMS
The PMS plan, PMPF plan and risk management file should feed each other, and the PMSR or PSUR schedule must match the device class.
How to Make Your File Easy to Review
- Follow the Annex II and Annex III order, or add a cross-reference table if your structure is different.
- Submit searchable, bookmarked PDFs with a clear document index.
- Give each document a unique number, version and approval date.
- Summarise long test reports and point to the exact page that supports each claim.
- Keep a change log so updates to the IVDR technical documentation after certification are easy to trace.
Keeping IVDR Technical Documentation Up to Date
Certification is not the end of the work. IVDR technical documentation is a living file that must reflect the current device, its current evidence and the current state of the art for as long as the device is on the market. Notified bodies check this during surveillance audits and unannounced audits, and competent authorities can request the file at any time.
Plan a regular review cycle and update the file whenever one of the following happens:
Design or intended purpose changes
Changes that could affect safety, performance or the intended purpose must be assessed, documented and, where required, reported to the notified body before they are implemented.
New post-market data
Complaints, trend reports, serious incidents, PMPF results and the PMSR or PSUR can change the benefit-risk conclusion, the risk file and the IFU.
New standards or common specifications
When a harmonised standard or common specification is revised, update the GSPR checklist and record a gap assessment for each affected requirement.
Supplier, site or process changes
New critical suppliers, manufacturing sites or processes must be reflected in the design and manufacturing information and in the risk management file.
Our IVDR consultants can run a periodic IVDR technical documentation review, so the file stays aligned with your PMS data and is ready for the next notified body audit.
IVDR Templates and Tools
Useful documents for teams building or updating IVDR technical documentation:
- IVDR technical documentation checklist (full Annex II and III table of contents)
- EU Declaration of Conformity (DoC) guide
- IVDR codes (PDF download)
- Risk analysis procedure & templates, useful for small and medium-sized manufacturers
- IVDR performance evaluation checklist
- IVDR CE marking cost calculator
Upgrading from IVDD?
Many self-declared IVDD devices move up to Class B, C or D under IVDR and need notified body review for the first time. Check the IVDR transition timeline and see our IVDR pricing.
Do you need an email containing full details about IVDR technical documentation within 2 minutes? Share your email below. Privacy Policy
Need Help With Your IVDR Technical Documentation?
Our IVDR team can prepare new technical documentation, remediate an IVDD file or respond to notified body findings.
- IVDR technical documentation preparation in STED format
- Technical documentation review and gap assessment
- GSPR checklists and standards mapping for Annex II and Annex III
- Performance evaluation, PMS and PMPF documents
- Notified body questions and deficiency support
Start your IVDR project
Share your device details and get a tailored proposal.
Request a QuoteContact UsIVDR Technical Documentation: Frequently Asked Questions
Is stability testing of the IVD device and of the specimen mandatory in the technical documentation?
Yes, both are mandatory. Accelerated stability data on its own is not sufficient to support shelf-life claims; real-time data is expected.
What should be included in the information supplied by the manufacturer?
All primary and secondary labelling, the instructions for use and any marketing material that makes claims. Every claim must be supported by evidence (IVDR Article 7).
Why include previous and similar generations of the device?
It gives the history of your device and helps the notified body understand how existing evidence relates to the current device and its claims.
What design information is needed?
Notified bodies need enough detail to understand the design and manufacture of the device, including design stages, manufacturing processes and sites.
What should the benefit-risk analysis and risk management section show?
These should be live documents that show continuous effort to understand and reduce risks while keeping the device compliant.
What analytical performance information is required?
Performance study reports and evaluations that explain how the analytical performance results support the intended purpose of the device.
What goes into the clinical performance and performance evaluation report?
The performance evaluation report (PER) presents evidence on scientific validity, analytical performance and clinical performance to demonstrate a favourable benefit-risk profile.
Who is responsible for IVDR technical documentation?
The manufacturer. It must draw up the technical documentation and keep it up to date (IVDR Article 10). If the manufacturer is outside the EU, its EU Authorised Representative must keep a copy available to competent authorities.
How long must IVDR technical documentation be kept?
For at least 10 years after the last device covered by the EU declaration of conformity has been placed on the market (IVDR Article 10(7)).
How is technical documentation for IVDR different from an IVDD technical file?
IVDR needs a structured Annex II and Annex III file, a GSPR checklist instead of the Essential Requirements, a full performance evaluation covering scientific validity, analytical and clinical performance, and PMS documents such as the PMPF plan, PMSR or PSUR. Most IVDD files need significant remediation. See our guide to the IVDD to IVDR transition.
Can one technical documentation file cover several devices?
Variants and configurations of the same device can share one file when they have the same intended purpose and each variant is clearly described. Devices with different intended purposes generally need separate files.
Does a Class A IVD need technical documentation?
Yes. Class A non-sterile devices are self-declared, but the manufacturer must still prepare full technical documentation under Annex II and Annex III, sign the EU declaration of conformity and keep the file up to date before and after placing the device on the market.
Updated | Author: Sobha Augustine, Founder, I3CGLOBAL | Reviewed by: Swathi Tummala, Senior Regulatory Consultant, I3CGLOBAL