
European Authorized Representative (EU REP) for Medical Devices and IVDs
A European Authorized Representative is the EU-based person or company that a manufacturer outside the EU must appoint, by written mandate, before placing a medical device or IVD on the EU market (MDR and IVDR Article 11). I3CGLOBAL acts as European Authorized Representative from offices in Germany and Portugal, and also handles your EUDAMED registration, SRN and UDI data.
What Is a European Authorized Representative?
A European Authorized Representative (also called EU REP, EC REP, EAR or, in British spelling, EU authorised representative) is a natural or legal person established in the EU that has received and accepted a written mandate from a non-EU manufacturer to act on its behalf for specific tasks under EU MDR 2017/745 or EU IVDR 2017/746. It is the official contact for EU authorities and shares legal liability with the manufacturer.
Key facts: mandatory for every non-EU manufacturer (MDR and IVDR Article 11) · one sole representative per generic device group · mandate valid only when accepted in writing · jointly and severally liable for defective devices · must have a PRRC at its disposal · named on the label and in the declaration of conformity.
| Question | Short answer |
|---|---|
| Legal basis | MDR and IVDR Article 11 (tasks), Article 12 (change of representative), Article 15(6) (PRRC) |
| Who must appoint one | Every manufacturer not established in an EU Member State, for every device class |
| When | Before the device is placed on the EU market |
| Where the CE mark applies | All 27 EU Member States plus Iceland, Liechtenstein and Norway, wherever the representative is located |
| Our addresses | Alsstr. 97, 41063 Mönchengladbach, Germany Rua Hermano Neves n° 18, Piso 3, E7, 1600-477 Lisboa, Portugal |
| Check a registration | Already working with us? Verify EAR status |
| Fees | Annual fee. See the EU Representative cost page or the fee calculator. |
Who Needs a European Authorized Representative?
If your company is not established in an EU Member State, you need a European Authorized Representative before you can sell any medical device or IVD in the EU. The device class does not matter: Class I and Class A devices need one too.
| Manufacturer located in | EU REP needed? |
|---|---|
| USA, Canada, China, India, Japan, South Korea and other non-EU countries | Yes, for every device placed on the EU market |
| United Kingdom | Yes. Since Brexit, UK manufacturers are non-EU manufacturers for the EU market. |
| Switzerland | Yes. The EU–Swiss mutual recognition agreement no longer covers medical devices. |
| Iceland, Liechtenstein, Norway (EEA) | No. EEA manufacturers are treated like EU manufacturers. |
| EU Member State | No. The manufacturer is the responsible EU actor itself. |
We represent manufacturers from Asia, North America and Europe, including China, India, South Korea, Japan, the USA, the UK and Vietnam.
What Does a European Authorized Representative Do?
Article 11(3) of the MDR and IVDR lists the minimum tasks the mandate must cover. A European Authorized Representative is not just an address on the label; these are legal duties. The European Commission guidance MDCG 2022-16 explains them in more detail.
- Verify documentsCheck that the EU declaration of conformity and technical documentation exist and that the right conformity assessment was done.
- Keep copiesHold the technical documentation, declaration of conformity and certificates for the period required by Article 10(8) (IVDR 10(7)).
- RegistrationMeet its own EUDAMED registration duties and verify the manufacturer’s UDI and device registration.
- Answer authoritiesGive competent authorities the information needed to show conformity, in an official EU language they accept.
- Samples and accessForward authority requests for samples or device access to the manufacturer and confirm they are met.
- Corrective actionsCooperate with authorities on preventive or corrective actions to remove or reduce risks.
- Complaints and incidentsImmediately pass complaints and suspected incident reports from users to the manufacturer.
- End the mandate if neededTerminate the mandate if the manufacturer acts against its MDR or IVDR obligations, and inform the authority.
The representative must also have a Person Responsible for Regulatory Compliance (PRRC) permanently available (Article 15(6)).
Is the European Authorized Representative Liable for Defective Devices?
Yes. If the manufacturer is not established in the EU and has not met its Article 10 obligations, the European Authorized Representative is legally liable for defective devices on the same basis as the manufacturer, jointly and severally (Article 11(5)). This is why a serious representative reviews your documentation before signing.
Manufacturer’s insurance
Under MDR Article 10(16) and IVDR Article 10(15), manufacturers must have measures for sufficient financial coverage of their potential liability, usually product liability insurance that remains valid while devices are on the EU market.
New Product Liability Directive
Directive (EU) 2024/2853 applies to products placed on the market from 9 December 2026 and replaces Directive 85/374/EEC. For non-EU manufacturers, the authorised representative and the importer can be held liable, and software is clearly covered.
How Do MDR and IVDR Rules for the EU REP Compare?
The IVDR copies the MDR rules for the authorised representative almost word for word. Only some article numbers differ.
| Topic | MDR 2017/745 | IVDR 2017/746 |
|---|---|---|
| Designation and tasks | Article 11 | Article 11 |
| Change of representative | Article 12 | Article 12 |
| Document retention period | Article 10(8) | Article 10(7) |
| Registration of actors | Article 31 | Article 28 |
| UDI and device registration checked | Articles 27 and 29 | Articles 24 and 26 |
| Financial coverage of manufacturer | Article 10(16) | Article 10(15) |
| PRRC at the representative | Article 15(6) | Article 15(6) |
What Do Our European Authorized Representative Services Include?
Unlike law firms or trading companies, I3CGLOBAL is a medical device regulatory consultancy, so your representative understands the technical documentation it signs for.
Documentation review
We review your declaration of conformity and technical documentation against MDR or IVDR and tell you about gaps.
Safe custody
We store the latest versions of your technical documentation and provide them to EU authorities on request.
EUDAMED and SRN
We complete your EUDAMED registration, SRN and UDI data.
PRRC in the EU
A qualified Person Responsible for Regulatory Compliance covers the representative’s regulatory duties.
Label address
Our name, address and EC REP symbol for your labels, packaging and IFU.
Vigilance link
Complaints go to your team; serious incidents, FSCAs and recalls are handled with the EU authorities without delay.
Authority liaison
Contact with competent authorities and Notified Bodies on your behalf.
Free sale certificates
Support for free sale certificates for export markets.
Data security
Customer documents are handled under our ISO/IEC 27001 certified information security system.
EU Representative in Germany or Portugal: Which Should You Choose?
Both offices carry out the full Article 11 mandate with the same regulatory team. Your CE mark is valid in every EU and EEA country whichever address you choose. The difference is commercial, not regulatory.
Credibility and commercial reach
- Best for Class IIb and III devices and higher-value products
- A German address many hospital buyers, tender panels and DACH distributors recognise
- Close to Düsseldorf, home of the MEDICA trade fair
- BfArM as your reference competent authority
Faster and more economical
- Best for start-ups, Class I and IIa devices, IVDs and cost-sensitive portfolios
- Lower annual fee than our German office, with the same mandate and EUDAMED support
- Leaner onboarding for tight launch dates
- Useful link to Portuguese-speaking markets such as Brazil
How Do You Appoint a European Authorized Representative?
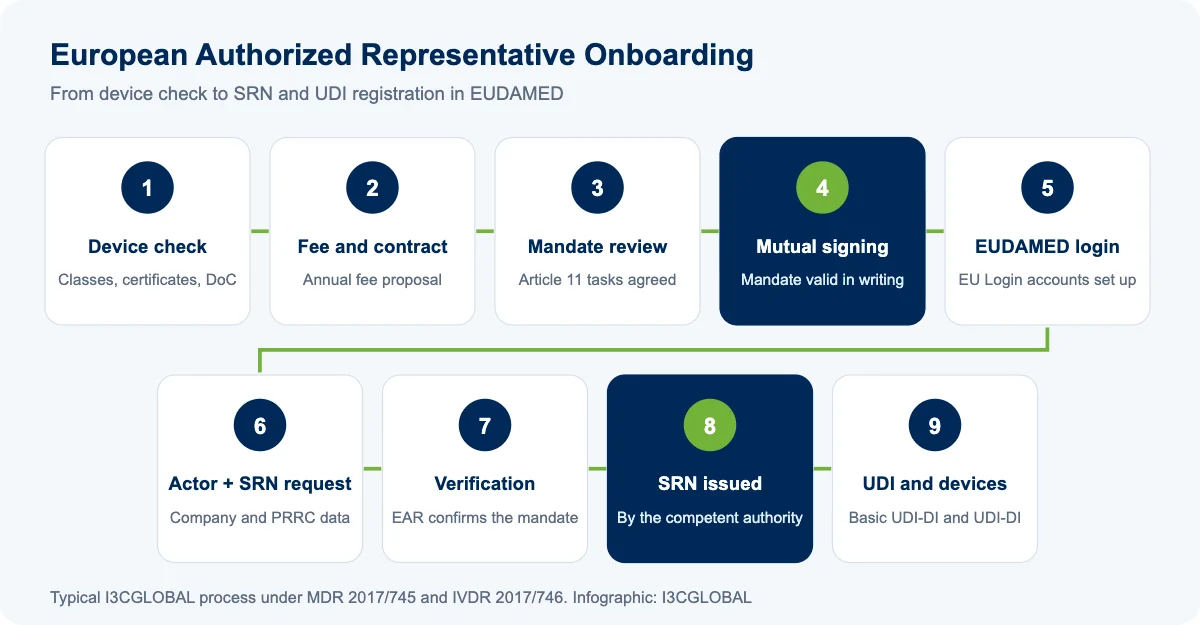
Appointment follows nine steps, from device check to SRN and UDI registration. The EU REP contract itself is usually signed within 4 working days of receiving your documents.
What Must the EU Representative Mandate Include?
The “agreement” and the “mandate” usually mean the same written contract. Under Article 11(2) and (3), it must:
- Be accepted in writing by the representative to be valid
- Cover at least all devices of the same generic device group
- List at least the eight tasks in Article 11(3)
- Not delegate the manufacturer’s core obligations listed in Article 11(4), such as the QMS, risk management and technical documentation
- Be provided to the competent authority on request
Our agreements are normally valid for five years unless terminated earlier under their terms. We provide our mandate template within 2 days of receiving the annual fee.
How Do You Change Your European Authorized Representative?
Under Article 12, a change of representative is set out in an agreement between the manufacturer, the outgoing representative (where practicable) and the incoming representative. The agreement must cover at least:
- The date the outgoing mandate ends and the date the incoming mandate starts
- The date until which the outgoing representative may still appear on information supplied, including promotional material
- The transfer of documents, including confidentiality and property rights
- The outgoing representative’s obligation, after the mandate ends, to forward complaints and suspected incident reports to the manufacturer or the incoming representative
Switching to I3CGLOBAL? We coordinate the transfer, update EUDAMED and plan the label change with you.
Must the EU REP Appear on the Device Label?
Yes. For manufacturers outside the EU, the name and registered address of the European Authorized Representative must be on the label (MDR and IVDR Annex I, Chapter III). It must also appear in the EU declaration of conformity.
- Use the EC REP symbol from EN ISO 15223-1 next to the representative’s name and address to avoid translations
- If the device label is too small, the information may go on the packaging or in the instructions for use, as allowed by Annex I
- Plan the label update before a change of representative, because old labels may only be used until the date agreed under Article 12
Do You Also Need a Representative for the UK, Switzerland or GPSR?
United Kingdom
To sell in Great Britain, non-UK manufacturers, including EU manufacturers, must appoint a UK Responsible Person and register devices with the MHRA.
Switzerland
Because the EU–Swiss mutual recognition agreement no longer covers medical devices and IVDs, manufacturers outside Switzerland need a Swiss authorised representative (CH REP).
GPSR
Medical devices and IVDs follow the MDR and IVDR, so they need an EU Authorized Representative under Article 11, not a GPSR responsible person. Other consumer products you sell may need one.
Get our EU REP services overview
Not ready to talk yet? We will email you our overview of the process, fees, timelines and MDR/IVDR scope. Privacy policy
Appoint I3CGLOBAL as Your European Authorized Representative
EU REP contract in 4 working days, EUDAMED registration in about 10 working days and a choice of German or Portuguese address, backed by a medical device regulatory team.
- MDR and IVDR mandate for all device classes
- EUDAMED actor, SRN, UDI and device registration
- PRRC, vigilance link and authority liaison
- Free sale certificates and UK Responsible Person services
Get your EU REP proposal
Tell us your device classes, number of devices and preferred office. We will send you a transparent fee proposal.
European Authorized Representative: Frequently Asked Questions
What is a European Authorized Representative?
A European Authorized Representative (EU REP, EC REP or EAR) is a person or company established in the EU that holds a written mandate from a non-EU medical device or IVD manufacturer to carry out specific regulatory tasks on its behalf under MDR or IVDR Article 11, and acts as the contact point for EU authorities.
Is a European Authorized Representative mandatory?
Yes. Under MDR and IVDR Article 11(1), a manufacturer not established in an EU Member State can place a device on the EU market only if it has designated a sole authorised representative. This applies to all device classes, including Class I and Class A.
What is the difference between EU REP, EC REP and EAR?
There is no legal difference. EU REP, EC REP, EAR and Authorised Representative all describe the same role. EC REP is also the name of the label symbol in EN ISO 15223-1 used next to the representative’s name and address.
Can a manufacturer have more than one EU Authorized Representative?
A manufacturer must have one sole authorised representative for each generic device group. Different generic device groups may be covered by different representatives, but most manufacturers use one representative for their whole portfolio.
Is the EU Authorized Representative liable for defective devices?
Yes. Under Article 11(5), if the manufacturer is not established in the EU and has not complied with its Article 10 obligations, the authorised representative is legally liable for defective devices on the same basis as, and jointly and severally with, the manufacturer.
Can a distributor or importer act as my EU Authorized Representative?
An importer or distributor may also be appointed as authorised representative if it accepts the written mandate and meets all Article 11 duties. Many manufacturers prefer an independent representative to keep regulatory control separate from commercial partners.
Does the EU REP need a PRRC?
Yes. Under Article 15(6), the authorised representative must have permanently and continuously at its disposal at least one Person Responsible for Regulatory Compliance with the required expertise.
Does the location of my EU REP limit where I can sell?
No. A CE marked device can be sold in all EU Member States and in Iceland, Liechtenstein and Norway, whichever EU country your authorised representative is in.
How long does it take to appoint I3CGLOBAL as EU REP?
Once we have your documents and the fee, the EU REP agreement is usually completed within 4 working days. EUDAMED actor registration and SRN usually follow within about 10 working days, depending on the competent authority.
How much does a European Authorized Representative cost?
I3CGLOBAL charges an annual fee that depends on the device class, the number of devices and the office you choose. See our EU Representative fee page or fee calculator, or request a proposal.
How do I change my European Authorized Representative?
Under Article 12, the manufacturer, the outgoing representative where practicable and the incoming representative sign an agreement covering the mandate end and start dates, how long the outgoing name may stay on labels, the transfer of documents and the forwarding of complaints and incident reports.
Do UK and Swiss manufacturers need an EU Authorized Representative?
Yes. Since Brexit, UK manufacturers are non-EU manufacturers and need an EU REP. Swiss manufacturers also need one because the EU–Swiss mutual recognition agreement no longer covers medical devices.
Is there a template for the EU Authorized Representative mandate?
There is no official EU template. The mandate must cover at least the tasks in MDR and IVDR Article 11(3), and the European Commission guidance MDCG 2022-16 explains what it should contain. I3CGLOBAL provides its mandate template within 2 days of receiving the annual fee.
Does I3CGLOBAL also act as UK Responsible Person?
Yes. In addition to EU representation from Germany and Portugal, I3CGLOBAL provides UK Responsible Person services for Great Britain. I3CGLOBAL does not act as Swiss authorised representative (CH REP).
EU Representative Guides and Related Services
Updated | Author: Sobha Augustine, Founder, I3CGLOBAL | Reviewed by: Joe Kumar, I3CGLOBAL