
Medical Device Testing
Medical device testing is the laboratory and bench evidence that shows a device is safe and performs as intended. I3CGLOBAL helps manufacturers plan the right tests, work with accredited laboratories and use the reports in EU MDR and US FDA submissions.
The basics
What Is Medical Device Testing?
Medical device testing is the set of laboratory, bench and simulated-use checks that verify a device meets its specifications and safety requirements. It includes biological, electrical, mechanical, sterility, packaging and performance work.
Test results support design verification and validation. They also feed the risk management file, the clinical evaluation and the technical documentation. Which tests apply depends on the materials, the contact with the body, the energy source, the sterility state and the claims.

Requirements
Why Is Medical Device Testing Required?
Regulators expect objective evidence, not statements. EU MDR 2017/745 asks for this evidence in its general safety and performance requirements and in the technical documentation. The US FDA expects performance data in submissions where it is relevant to the device.
| Source | What It Asks For | Effect on Testing |
|---|---|---|
| EU MDR Annex I | General safety and performance requirements, including chemical, biological, electrical, mechanical and infection risks. | Each applicable requirement needs supporting evidence. |
| EU MDR Annex II, Section 6.1 | Pre-clinical data, such as results of engineering, laboratory, simulated use and animal tests. | Test reports, or a justification for not testing, go into the MDR technical file. |
| EU IVDR | Analytical performance and safety evidence for IVDs. | Evidence goes into the IVDR technical documentation. |
| US FDA | Performance and biocompatibility data, where applicable, in premarket submissions. | Testing supports a FDA 510(k) or other submission. |
| ISO 13485 | Design verification, validation and monitoring of processes. | Test records form part of the quality system. |
Testing also protects the business. Finding a design weakness before submission costs less than a failed test, a Notified Body question or a field safety corrective action.
Test types
What Are the 7 Main Types of Medical Device Testing?
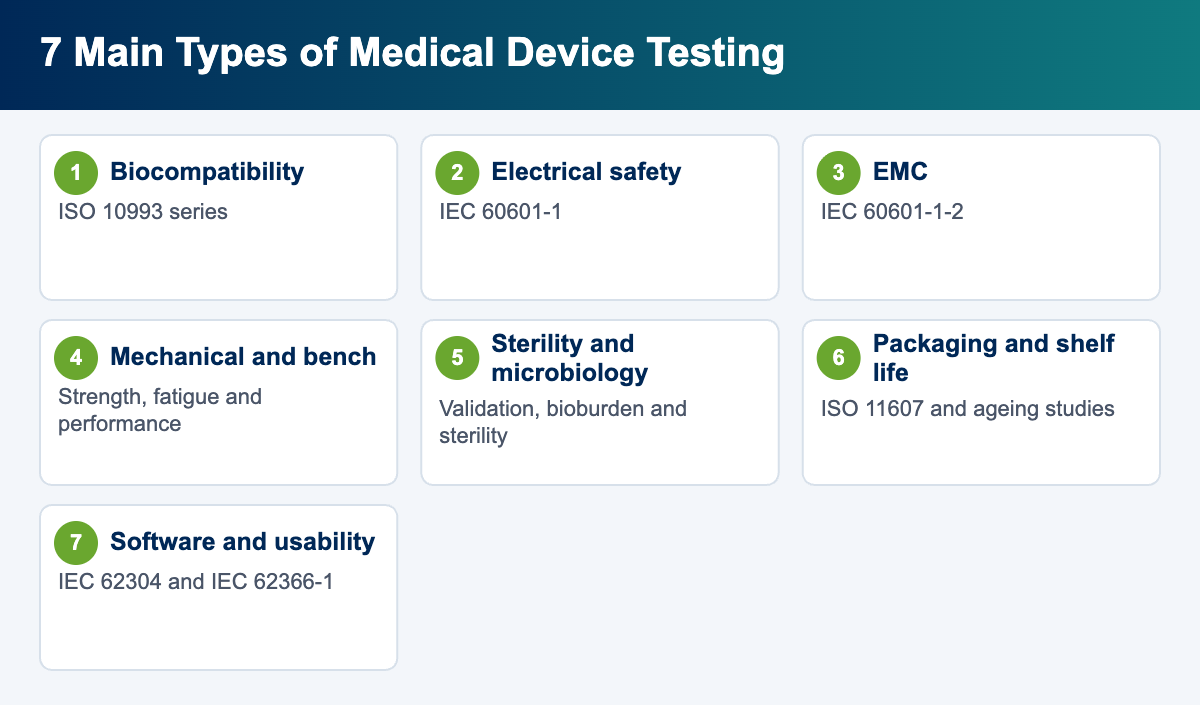
Most test programmes draw on these seven groups. A simple non-sterile device may need only a few, while an active implant may need nearly all of them.
1. Biocompatibility
Biocompatibility testing checks how materials and the final device interact with the body under ISO 10993.
2. Electrical safety
Electrical safety testing checks protection against shock and other hazards under IEC 60601-1.
3. EMC
Electromagnetic compatibility testing under IEC 60601-1-2 checks emissions and immunity.
4. Mechanical and bench
Strength, fatigue, wear, dimensional and durability tests show that the device withstands intended use.
5. Sterility and microbiology
Sterilisation validation, bioburden, sterility and endotoxin tests apply to sterile or low-bioburden devices.
6. Packaging and shelf life
Packaging, transport and ageing tests show that the device stays safe up to its stated expiry date.
7. Software and usability
Software verification under IEC 62304 and usability work under IEC 62366-1 cover software and user interface risks.
Material characterisation and chemical analysis often come first, as they can reduce later testing. The results of biological work are brought together in a biological evaluation report.
Test selection
How Do You Choose the Right Tests?
Test selection should come from the device and its risks, not from a generic list. Use these questions to scope the medical device testing plan.
| Question | Why It Matters | Likely Testing |
|---|---|---|
| Does it touch the body? | Contact type and duration set the biological evaluation. | Chemical characterisation and ISO 10993 endpoints |
| Does it use electricity? | Shock, heat and interference hazards apply. | IEC 60601-1 and IEC 60601-1-2 |
| Is it supplied sterile? | Sterility must be validated and maintained. | Sterilisation validation and packaging tests |
| Is it reusable? | Cleaning and reprocessing must be effective. | Reprocessing validation |
| Does it bear load or move? | Failure could cause harm. | Mechanical and fatigue testing |
| Does it have software? | Software errors can create hazards. | Software verification and validation |
| Is it used by lay persons? | Use errors are more likely. | Usability engineering and home-use standards |
Link every test to a requirement and a risk. Risk management should explain why a test is needed, and also why a test is not needed. A documented rationale is often as valuable as a report.
Standards
Which Standards Apply to Device Testing?
Standards are normally voluntary, but they are the most common way to show conformity. Under EU law, applying a harmonised standard gives a presumption of conformity with the requirements it covers. Check the current edition before you commit to a test.
| Area | Common Standards |
|---|---|
| Biological evaluation | ISO 10993 series |
| Electrical safety | IEC 60601-1 and relevant particular standards |
| EMC | IEC 60601-1-2 |
| Laboratory and IVD equipment | IEC 61010 series |
| Sterilisation | ISO 11135, ISO 11137 and ISO 17665 |
| Microbiology | ISO 11737 series |
| Sterile packaging | ISO 11607 series |
| Software | IEC 62304 |
| Usability | IEC 62366-1 |
| Risk management | ISO 14971 |
| Laboratory competence | ISO/IEC 17025 |
Standards change. For example, Implementing Decision (EU) 2026/1231 updated the harmonised standards list under the MDR in June 2026, so older reports may need a gap review rather than automatic acceptance.
Step by step
How Do You Plan a Test Programme?
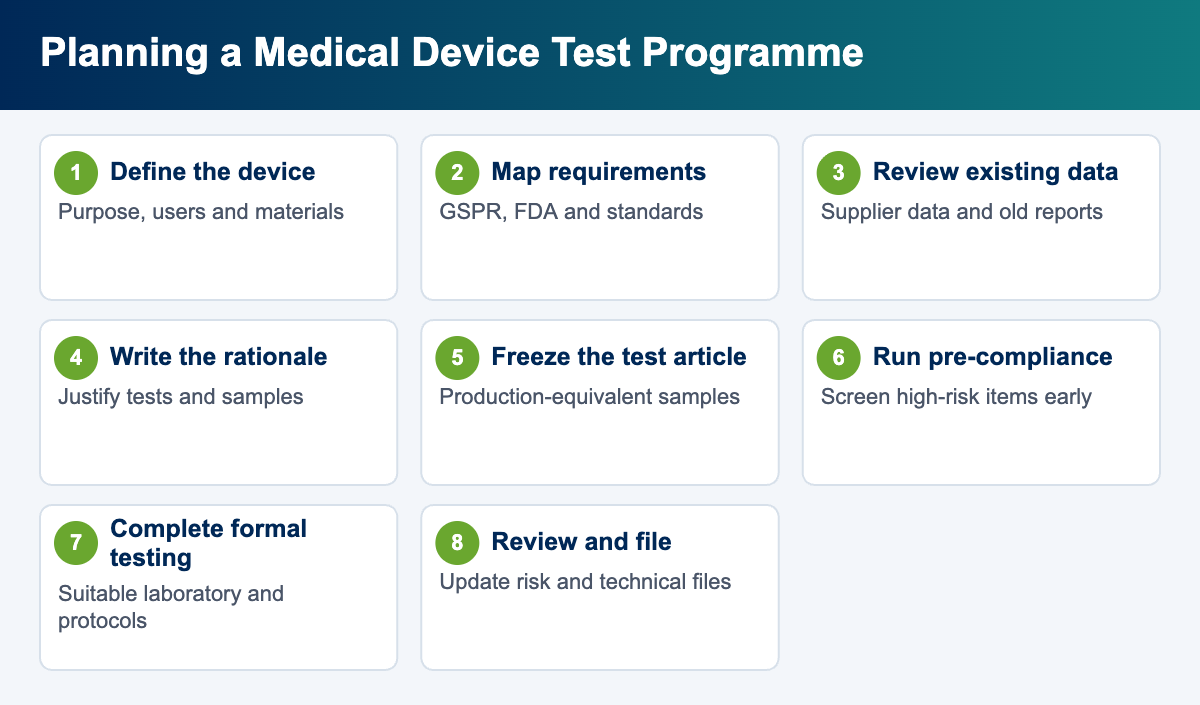
A good plan avoids repeat testing and delays. These steps fit most projects.
- Define the deviceRecord the intended purpose, users, materials, variants and accessories.
- Map the requirementsList applicable GSPR, FDA expectations and standards for your markets.
- Review existing dataCheck supplier data, earlier reports and literature before you buy new tests.
- Write the rationaleJustify each test, sample size and worst-case configuration.
- Freeze the test articleTest production-equivalent samples, because later changes can invalidate the results.
- Run pre-compliance checksScreen high-risk electrical, EMC or mechanical items early.
- Complete formal testingUse a suitable laboratory and approved protocols.
- Review and file reportsAssess any failures, update risk files and add the reports to the technical file.
Allow time for sample production, laboratory queues and retests. Long studies, such as chronic biological tests or real-time ageing, need particular care in your project timeline.
Laboratories
How Do You Choose a Medical Device Testing Laboratory?
Not every test report will satisfy a Notified Body or the FDA. Check the laboratory before you send samples, and ask these questions:
- Is the laboratory accredited to ISO/IEC 17025 for the exact test and standard?
- Does the scope include the current edition of the standard?
- Are animal studies performed under Good Laboratory Practice, where this is expected?
- Does the team have experience with your device type?
- Can it agree a protocol before testing starts?
- Will the report identify the test article, method, deviations and results?
- If you plan an FDA submission, does the laboratory take part in the FDA ASCA programme for relevant tests?
Documentation
What Should Protocols and Test Reports Include?
Test reports are read by people who did not watch the test. Each one should allow a reviewer to confirm what was tested, how and with what result.
| Document | Key Content |
|---|---|
| Test rationale | Reason for each test or waiver, linked to risks and requirements. |
| Protocol | Method, standard and edition, sample size, acceptance criteria and test article. |
| Test report | Device identity, batch, conditions, results, deviations, conclusion and signatures. |
| Summary | Results across all tests and how they meet each requirement. |
| Traceability | Links to GSPR checklist, risk file and design verification records. |
Keep results consistent with your labelling. For example, a claim for a seven-day wear time needs evidence that covers seven days of contact.
Common gaps
What Testing Gaps Should You Look For?
- Test article differs from the final production device
- Reports use a superseded standard, with no gap assessment
- A generic test list, with no device-specific rationale
- Sterilisation or packaging changes made after the testing
- Worst-case variant not justified
- Supplier certificates used in place of final device evidence
- Failed or repeated tests not explained
- Lab scope does not cover the method used
- Test results not linked to the risk file or GSPR checklist
Review these points before you submit your technical documentation. It is usually easier to close them early than during a Notified Body review.
Examples
Which Medical Device Testing Applies by Device Type?
These examples show how the test mix changes with the device. They are starting points only, since your own risk analysis and intended purpose decide what is needed.
| Device Type | Typical Testing Focus |
|---|---|
| Sterile single-use device | Biological evaluation, sterilisation validation, packaging, shelf life and functional performance. |
| Implant | Biological evaluation including implantation, mechanical fatigue, corrosion or wear, sterility and packaging. |
| Powered monitor or therapy device | Electrical safety, EMC, usability, software, performance and alarm testing where relevant. |
| Reusable instrument | Cleaning and reprocessing validation, durability, biological evaluation and mechanical performance. |
| Wearable device | Skin contact evaluation, electrical safety, EMC, battery safety, wireless function and ingress protection. |
| Medical device software | Software verification and validation, cybersecurity, usability and clinical or analytical performance evidence. |
| Laboratory or IVD equipment | IEC 61010 electrical safety, EMC, and analytical performance for the intended use. |
Accessories, single-use parts and replaceable components should be assessed too. A reusable monitor may need electrical testing, while its sensor also needs a biological evaluation.
Performance
What Does Mechanical and Bench Testing Cover?
Bench testing shows that a device performs to specification and withstands intended use. It is often the main evidence for lower-risk devices, and it supports clinical evaluation for higher-risk devices.
- Tensile, compression, bending and burst strength
- Fatigue and cycle life
- Dimensional and tolerance checks
- Connector and leak integrity
- Accuracy, precision and repeatability
- Simulated use and transport conditions
Set acceptance criteria in advance and base them on the requirements, standards or a comparison with benchmark devices. Statistical sample sizes should link to the risk of failure.
Sterility and packaging
What Testing Do Sterile Devices Need?
Sterility cannot be shown by testing the finished product alone. It depends on a validated sterilisation process, a controlled environment and a packaging system that maintains the sterile barrier.
| Evidence | Purpose | Common Standard |
|---|---|---|
| Sterilisation validation | Shows the process consistently achieves the required sterility assurance level. | ISO 11135, ISO 11137 or ISO 17665 |
| Bioburden | Measures microorganisms before sterilisation. | ISO 11737-1 |
| Tests of sterility | Used as part of validation and specific process checks. | ISO 11737-2 |
| Residues | Assesses residues, such as those from ethylene oxide. | ISO 10993-7 |
| Sterile barrier validation | Shows packaging protects the device up to use. | ISO 11607 series |
| Shelf life | Shows the device and packaging remain acceptable over the stated life. | Accelerated and real-time ageing studies |
Accelerated ageing can support an initial expiry date, but real-time ageing should follow to confirm it. Reusable devices need validated cleaning, disinfection and sterilisation instructions instead.
Market differences
How Do EU and FDA Testing Expectations Differ?
The same test report can often support both markets, but you should plan for the differences at the start of the project.
| Topic | EU MDR | US FDA |
|---|---|---|
| Main reviewer | Notified Body for most devices above Class I. | FDA, usually through a 510(k), De Novo or PMA. |
| Standards | Harmonised standards can give a presumption of conformity. | Recognised consensus standards can support a declaration of conformity. |
| Evidence focus | General safety and performance requirements. | Substantial equivalence or safety and effectiveness, depending on the pathway. |
| Biological safety | ISO 10993 series within risk management. | ISO 10993-1 as applied in FDA guidance. |
| Comparison | Equivalence and clinical evaluation have their own rules. | Performance is often compared with a predicate in a 510(k). |
Our US FDA consultants and EU teams can help you build one plan that covers both markets, which can reduce repeat testing.
Planning
What Affects the Cost and Timing of Testing?
There is no standard price for a test programme. These factors have the greatest effect on the budget and the schedule:
Device complexity
More functions, materials and variants mean more testing.
Contact duration
Long-term contact generally needs more biological evidence.
Sample availability
Production-equivalent samples take time to make.
Pre-compliance work
Early checks cost time now, but can avoid failed formal tests.
Laboratory capacity
Popular laboratories may have long queues.
Retesting
Design changes after testing can add cost and delay.
Request quotes that state the method, standard edition, sample numbers and deliverables, so you can compare them fairly. The lowest quote may not cover what a reviewer expects.
Change control
When Must Testing Be Repeated?
Medical device testing does not end at approval. Your change control process should ask whether each change affects existing evidence. Common triggers include:
- A new material, colourant, adhesive or supplier
- A change to a critical electrical component or firmware
- A new sterilisation method or site
- New packaging or a longer shelf life
- A new intended use, patient group or environment
- An updated standard, with a relevant technical change
- Complaints or post-market data that question a test assumption
Record the assessment, even if no retesting is needed. Significant changes may also need notification to your Notified Body before they are implemented.
Our services
How Does I3CGLOBAL Help With Testing?
I3CGLOBAL provides regulatory consulting and arranges testing through accredited laboratories. We support manufacturers in these areas:
Test strategy
Identify the tests your device needs for EU, UK and US submissions.
Biological evaluation
Plan testing and prepare the biological evaluation plan and report.
Electrical safety
Plan IEC 60601-1 and EMC work and review the reports.
Laboratory coordination
Support sample preparation, protocols and communication with the laboratory.
Report review
Check reports for gaps before they go into your technical file.
Regulatory integration
Link results to the risk file, GSPR checklist and clinical evaluation.
Work with I3CGLOBAL
Plan Your Medical Device Testing
Tell us about your device and target markets, and our team will suggest a practical test plan.
- Test strategy and gap review
- Laboratory coordination
- Biological and electrical safety evidence
- Report review for submissions
Request your proposal
Share your device details, materials and target markets, and we will prepare a proposal.
FAQs
Medical Device Testing: Frequently Asked Questions
What is medical device testing?
Medical device testing is the laboratory, bench and simulated-use work that shows a device meets its specifications and safety requirements. It commonly includes biocompatibility, electrical safety, EMC, mechanical, sterility, packaging and software testing.
Is medical device testing mandatory?
Regulators require evidence of safety and performance, and testing is the usual way to provide it. A specific test may not be needed if a documented rationale, existing data or a change assessment justifies it.
What tests are needed for CE marking?
This depends on the device. Common areas are biological evaluation, electrical safety and EMC for active devices, sterility and packaging for sterile devices, mechanical performance and software verification.
What is the difference between verification and validation testing?
Verification checks that design outputs meet design inputs. Validation checks that the finished device meets user needs and its intended use under defined conditions.
Do test reports need to come from an accredited laboratory?
Check the expectations of your Notified Body or regulator. ISO/IEC 17025 accreditation for the exact test scope is widely expected, and it gives reviewers greater confidence in the results.
Can supplier data replace device testing?
Sometimes it can support your evidence, but it rarely replaces evaluation of the final device. Processing, assembly, sterilisation and packaging can change the final product.
Do I need to repeat testing after a design change?
Assess each change. A change to material, supplier, process, software, sterilisation or packaging may need a new test, while other changes can be justified without retesting. Document the rationale.
How long does medical device testing take?
Timing varies with the tests, samples and laboratory workload. Short tests may take weeks, while chronic biological studies and real-time ageing can take much longer, so plan early.
Are the same tests accepted in the EU and USA?
Many tests overlap, as both markets use ISO and IEC standards, but expectations differ. Check FDA recognised standards and guidance, and the harmonised standards and Notified Body expectations in the EU.
Can I3CGLOBAL arrange testing?
Yes. I3CGLOBAL provides testing services through accredited laboratories and helps with test planning, protocols and report review for EU MDR and US FDA submissions.