
Biological Evaluation Report and Biological Evaluation of Medical Devices
A biological evaluation report (BER) records how the biological safety of a medical device was assessed and what was concluded. I3CGLOBAL helps manufacturers plan the biological evaluation of medical devices and prepare reports for EU MDR, UK and US FDA submissions.
The basics
What Is a Biological Evaluation Report?
A biological evaluation report is the document that brings together all biological safety evidence for a device. It summarises tests performed, justifications for tests not performed, chemistry, literature and other existing data, and it gives a conclusion on biological risk.
It is not just a list of test reports. A good report explains how the data apply to the final device, its intended contact and its users, and it confirms the outcome of the biological risk assessment.
Requirements
Why Is a Biological Evaluation Report Required?
Regulators require biological safety to be demonstrated where a device contacts the body. A report is the usual way to present this evidence.
| Source | Requirement | Role of the Report |
|---|---|---|
| EU MDR Annex I, Section 10 | Chemical, physical and biological properties, including compatibility with tissues, cells and body fluids. | Shows how biological requirements are met. |
| EU MDR Annex II, Section 6.1 | Pre-clinical data, including biocompatibility, in the technical documentation. | Part of the MDR technical file. |
| EN ISO 10993-1:2025 | Biological evaluation within a risk management process. | Follows the process and records the conclusions. |
| ISO 14971 | Risk management. | Links biological hazards to overall risk evaluation. |
| US FDA | Biocompatibility expectations in premarket submissions. | Supports a 510(k) submission. |
The process
What Is the Biological Evaluation of Medical Devices?
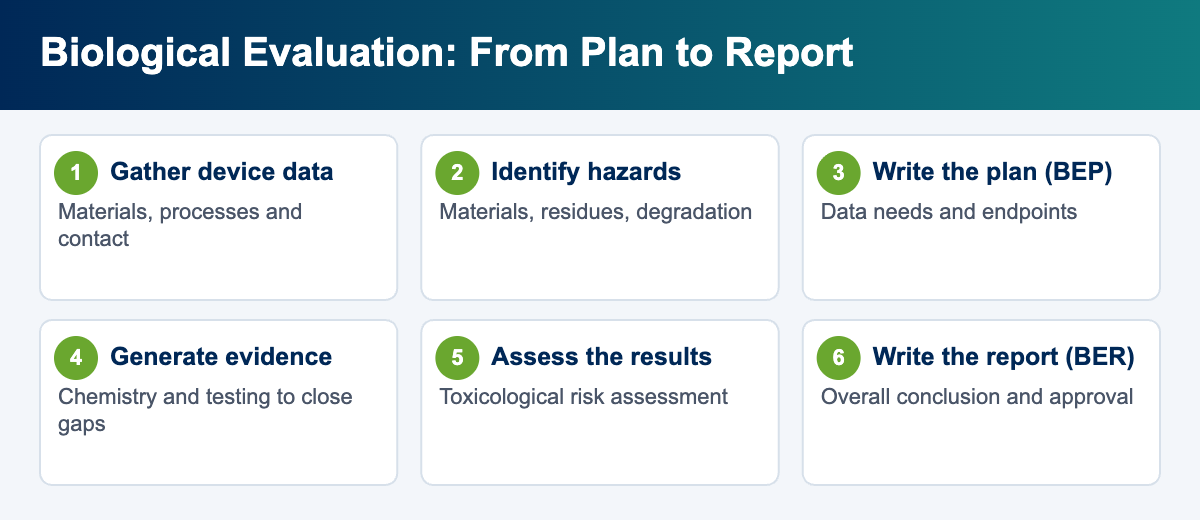
The biological evaluation of medical devices is a structured, risk-based assessment under the ISO 10993 series. It identifies biological hazards, estimates exposure and decides what evidence is needed to show that risks are acceptable.
It is wider than biocompatibility testing. It may use material data, chemical characterisation, literature, existing test results and clinical experience. New testing is used to close a gap, not as an automatic first step.
Key documents
What Is the Difference Between a BEP and a BER?
A biological evaluation plan (BEP) is prepared before the evaluation is carried out. The report is prepared afterwards. Both should form part of your risk management activity.
| Point | Biological Evaluation Plan | Biological Evaluation Report |
|---|---|---|
| Purpose | Sets out how the evaluation will be done. | Records what was done and what was concluded. |
| Timing | At the start, and before testing. | After data review and testing. |
| Content | Device data, contact, hazards, data sources, proposed endpoints and tests. | Results, data assessment, justifications, risk conclusions and overall conclusion. |
| Reviewer use | Shows that the strategy was planned. | Shows that biological safety was demonstrated. |
| Updates | When the strategy changes. | When new information or changes affect the conclusion. |
The plan should describe arrangements for gathering literature, reviewing and approving the plan in design control, approving additional testing and reviewing the final biological risk assessment.
Document structure
What Are the 9 Essential Contents of a Biological Evaluation Report?
The structure can vary, but reviewers normally look for these nine elements:
1. Scope and device description
Device, variants, intended use, users and packaging.
2. Contact and exposure
Contact type, duration, frequency and foreseeable misuse.
3. Materials and processing
All contact materials, additives, residues, sterilisation and manufacturing.
4. Existing data
Literature, supplier data, earlier tests and clinical or post-market data.
5. Chemical characterisation
Analytical results and the rationale, where used.
6. Endpoint assessment
How each biological endpoint is addressed.
7. Test results
Summary of reports, deviations and result interpretation.
8. Risk assessment
Toxicological assessment and link to the risk management file.
9. Conclusion and competence
Overall conclusion, evaluator qualifications, approval and any further actions.
Justifications for tests not performed deserve special care. A short statement is rarely enough, so explain which data support the decision and why they apply to your device.
Step by step
How Do You Prepare a Biological Evaluation Report?
- Gather device informationCollect drawings, material lists, process data and intended use.
- Categorise the contactDefine the nature and duration of contact under the current standard.
- Review existing informationSearch literature and review earlier reports and post-market data.
- Identify hazardsIdentify biological hazards from materials, processing, degradation and residues.
- Write the planSet out data needs, endpoints and any proposed tests.
- Generate new evidenceComplete chemistry and testing to close the gaps.
- Assess the resultsInterpret results and complete toxicological assessment, where needed.
- Conclude and approveFinalise the report with qualified reviewers and link it to the risk file.
For a quicker start, our biological evaluation procedure and templates are available as editable documents, or our team can prepare the documents for you.
Evaluation inputs
Which Factors Should the Evaluation Consider?
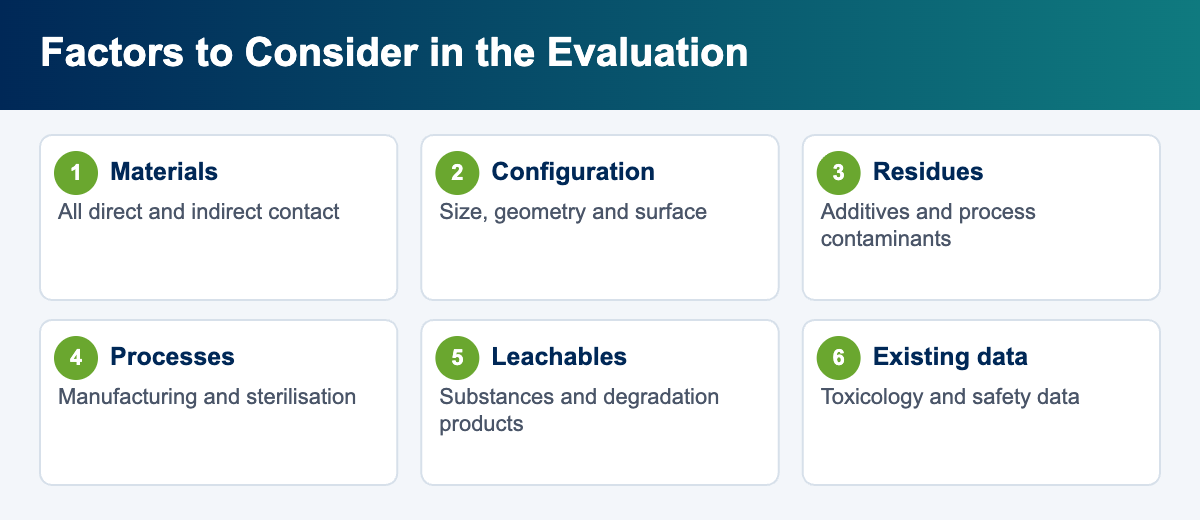
- All direct and indirect contact materials
- Device configuration, such as size, geometry and surface properties
- Additives, process contaminants and residues
- Manufacturing, cleaning and sterilisation processes
- Packaging that may contact the device
- Leachable substances and degradation products
- Interaction of components in the final product
- Physical and chemical characteristics of the device
- Local and systemic effects
- Existing toxicology and other safety data
Latest standard
How Does ISO 10993-1:2025 Affect Your Report?
ISO 10993-1:2025 was published in November 2025, and EN ISO 10993-1:2025 is reported as harmonised under the MDR in June 2026. Older reports should have a gap assessment. Key points include:
| Change | What to Check in Your Report |
|---|---|
| Integration with ISO 14971 | Biological risk estimation is clear and linked to the risk file. |
| Four contact-based tables | Endpoints are re-mapped against the new tables. |
| Exposure based on contact days | Repeated use has been assessed correctly. |
| Reasonably foreseeable misuse | Misuse is addressed from the planning stage. |
| Biological equivalence | Any equivalence claim meets the requirements of the new edition. |
| Evaluator expertise | Qualifications are shown for those who performed the evaluation. |
| Lifecycle evaluation | Post-market information is reviewed and used in updates. |
Maintenance
When Should You Update the Report?
A biological evaluation report is a living document. Review it periodically, and also after any event that could affect biological safety:
- Change in material, formulation or supplier
- Change in manufacturing, cleaning or sterilisation process
- Change in packaging or storage conditions
- New intended use, patient group or contact duration
- New toxicological information about a material or substance
- Adverse events, complaints or post-market findings
- A relevant change in standards or regulatory requirements
Common gaps
What Gaps Do Reviewers Often Find?
- The report lists tests, but gives no overall risk conclusion
- Unsupported justifications for tests not performed
- Test article not linked to the final device
- Missing indirect contact materials, such as fluid paths
- Contact duration understated for repeated use
- Supplier certificates used without further evaluation
- Chemistry results with no toxicological assessment
- Report not linked to the risk management file
- No evidence of evaluator competence
- No update after changes or the new ISO 10993-1 edition
Evidence
How Are Biological Endpoints Addressed in the Report?
The report should show how each relevant endpoint has been addressed. There may be more than one way to do this, and the choice should be justified.
| Evidence Route | When It May Be Used | What to Show |
|---|---|---|
| New test | A data gap remains and testing is the suitable way to close it. | Protocol, sample, result and interpretation. |
| Earlier test | An earlier report covers the same or a justified similar device. | Comparison of materials, process and contact. |
| Chemical characterisation | Chemistry and toxicology can address the risk. | Analytical data, exposure estimate and risk assessment. |
| Literature | Published data apply to the material and its use. | Search method, relevance and quality of the data. |
| Clinical or post-market data | Long use supports the conclusion, with no changes that matter. | Device identity, exposure, complaints and limits of the data. |
| Justification | The endpoint is not relevant or the risk is negligible. | A scientific rationale linked to the device. |
Keep the logic visible. A reviewer should be able to follow each endpoint from hazard to evidence to conclusion, without reading every source report.
Equivalence
Can You Use Biological Equivalence?
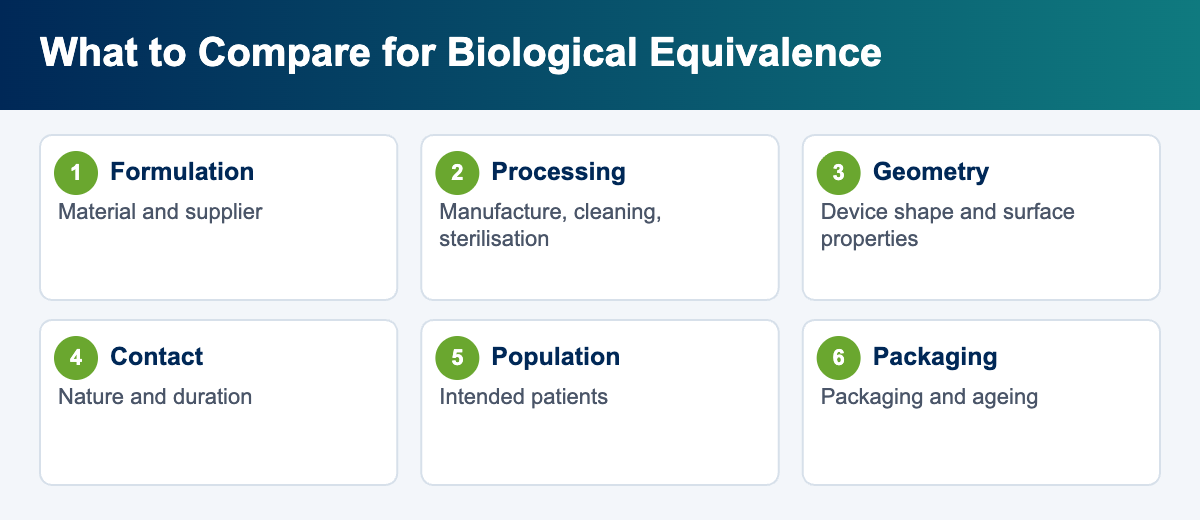
ISO 10993-1:2025 includes a dedicated section on biological equivalence. It can reduce new testing, but the comparison must be thorough. Consider these points:
- Formulation and material supplier
- Manufacturing, cleaning and sterilisation
- Device geometry and surface properties
- Nature and duration of contact
- Patient population
- Packaging and ageing
Do not rely on a statement that a material is medical grade. Medical grade is a supplier description, not proof of biological safety for your device.
Risk management
How Does the Report Link to Risk Management?
Biological evaluation is a risk management activity. The plan and report should connect directly with your medical device risk management file under ISO 14971.
| ISO 14971 Stage | Biological Evaluation Input |
|---|---|
| Risk management plan | Biological evaluation plan and acceptance criteria. |
| Risk analysis | Biological hazards from materials, residues and degradation, plus exposure. |
| Risk evaluation | Toxicological and biological conclusions compared with criteria. |
| Risk control | Material changes, process controls or information for safety. |
| Overall residual risk | Conclusion on biological safety for the final device. |
| Post-production | Complaints, adverse events and new toxicology data. |
If a biological risk needs a warning or a contraindication, make sure that it also appears in the label and instructions for use.
Notified Body review
What Do Notified Bodies Look For?
Reviewers want to see a clear scientific argument. Before you submit, check that your report answers these questions:
- Is the device in the report the same one in the technical file?
- Are all contact materials listed, with their manufacturing residues?
- Is the contact type and duration correct, including repeat use?
- Is the latest applicable edition of each standard addressed?
- Are test articles traceable to production?
- Are unusual results explained, rather than ignored?
- Do justifications rely on evidence, not on general statements?
- Is the conclusion consistent with the risk file, the clinical evaluation and the instructions?
- Are the authors and reviewers suitably qualified?
Choosing an approach
Should You Use a Template or Expert Support?
The right choice depends on your device and internal experience. A template gives structure, but it cannot make the scientific judgement for you.
| Option | Best For | Consider |
|---|---|---|
| Templates | Teams with biological evaluation experience and simpler devices. | You must still supply the rationale and data. |
| Review by an expert | Teams that have a draft or older report. | A gap assessment can identify priority work. |
| Full preparation | Complex devices, or teams without toxicology support. | Allow time to collect material and process data. |
Whichever route you choose, start by collecting complete material information from suppliers. Missing composition data is one of the most common causes of delay.
Getting started
What Information Is Needed Before You Start?
A complete data pack reduces rework. Collect as much of this information as you can before starting the evaluation:
Device data
Drawings, variants, accessories and intended use.
Material data
Composition, supplier declarations and safety data.
Process data
Moulding, cleaning, adhesives, coatings and sterilisation.
Existing reports
Biocompatibility, chemical and ageing reports.
Post-market data
Complaints, vigilance data and any similar device data.
Risk documents
Risk management plan, hazard analysis and instructions for use.
Our services
How Does I3CGLOBAL Help?
I3CGLOBAL’s regulatory professionals support biological evaluation projects for any risk class of device, for MDR CE marking and US FDA 510(k) submissions.
Plan preparation
Write the biological evaluation plan for your device.
Report preparation
Write or update the report, including justifications.
Gap assessment
Review older documents against ISO 10993-1:2025.
Testing support
Arrange testing through accredited laboratories, as needed.
Toxicology coordination
Coordinate chemical characterisation and risk assessment.
Templates
Provide editable procedure, plan and report templates.
Work with I3CGLOBAL
Get Your Biological Evaluation Report Prepared
Tell us about your device and existing data, and our team will suggest the next step.
- Biological evaluation plan and report
- ISO 10993-1:2025 gap assessment
- Testing through accredited laboratories
- Support for CE marking and FDA 510(k)
Request your proposal
Share your device details, materials and existing reports, and we will prepare a proposal.
FAQs
Biological Evaluation Report: Frequently Asked Questions
What is a biological evaluation report?
A biological evaluation report is a document that summarises all biological safety evidence for a medical device, including tests, justifications, chemistry and literature. It concludes whether the biological risks are acceptable.
What is biological evaluation of medical devices?
It is a structured, risk-based assessment under the ISO 10993 series. It identifies biological hazards, estimates exposure and determines what evidence is needed to show that the device is biologically safe.
What is the difference between a BEP and a BER?
A biological evaluation plan sets out how the evaluation will be done before it begins. A biological evaluation report records the data, assessment and conclusions after the work is completed.
Is a biological evaluation report mandatory?
Biological safety evidence is required for devices that contact the body, and ISO 10993-1 describes a documented evaluation. A report is the usual way to present it for a Notified Body or regulator.
Who should write a biological evaluation report?
People with suitable knowledge and experience should prepare it, often with toxicology and regulatory input. ISO 10993-1:2025 expects evaluators to demonstrate appropriate expertise.
Can a report replace biocompatibility testing?
A report does not replace needed data, but a justified evaluation may reduce or avoid some tests if existing data, chemistry or other evidence adequately address the endpoint.
Does my report need updating for ISO 10993-1:2025?
Carry out a gap assessment. Changes to exposure, misuse, endpoint tables and risk integration may affect your report. Ask your Notified Body how it expects you to transition.
Is a biological evaluation report needed for FDA 510(k)?
FDA expects biocompatibility information where applicable. Many manufacturers use a biological evaluation report or similar summary to organise the rationale, but check the current FDA guidance.
When should a biological evaluation report be updated?
Update it after changes to materials, suppliers, processes, sterilisation, packaging or intended use, and after relevant post-market findings or standard changes. Review it periodically too.
Does I3CGLOBAL provide biological evaluation report templates?
Yes. I3CGLOBAL offers editable biological evaluation procedure, plan and report templates, and our team can also prepare the documents for your device.