
FDA QMSR Consultants for 21 CFR Part 820 Compliance
I3CGLOBAL is a regulatory consulting company that helps medical device manufacturers meet the FDA Quality Management System Regulation (QMSR), the current version of 21 CFR Part 820. The QMSR took effect on February 2, 2026 and incorporates ISO 13485:2016 by reference. Our 21 CFR 820 consultants support manufacturers in the United States and those exporting to it, from gap analysis and QMS documentation to internal audits, inspection readiness and FDA Form 483 responses.
The basics
What Is the FDA QMSR (21 CFR 820)?
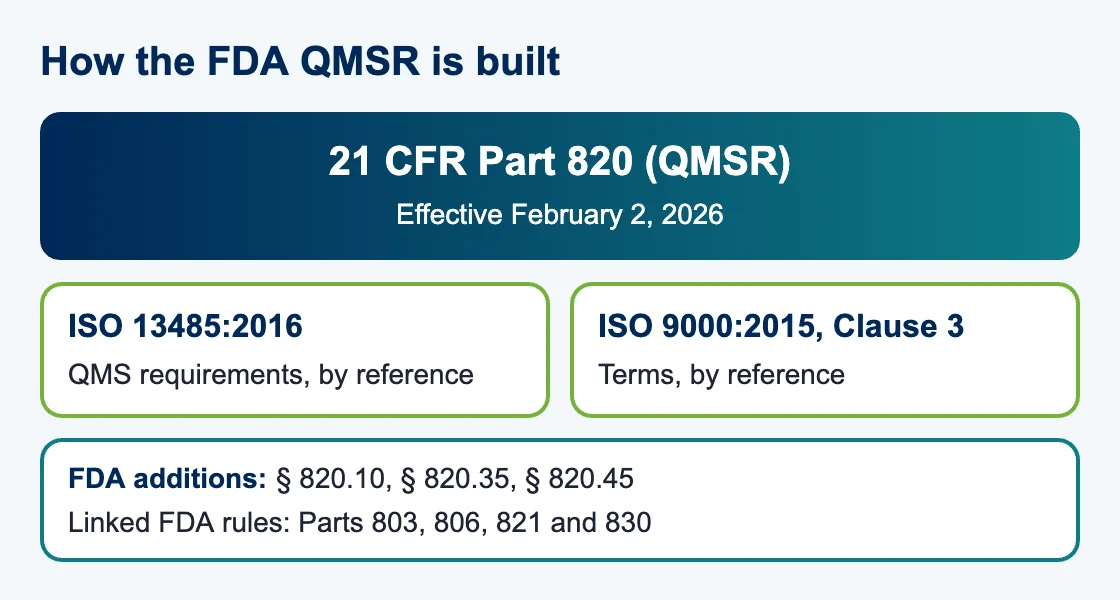
The Quality Management System Regulation (QMSR) is the FDA regulation at 21 CFR Part 820. It sets current good manufacturing practice (CGMP) requirements for finished medical devices. It replaced the former Quality System Regulation (QSR) on February 2, 2026.
The regulation covers the methods, facilities and controls used to design, manufacture, package, label, store, install and service finished devices. Its aim is that devices are safe and effective and comply with the Federal Food, Drug, and Cosmetic Act (FD&C Act).
The FDA QMSR does this by incorporating two standards by reference:
- ISO 13485:2016, the international standard for medical device quality management systems.
- Clause 3 of ISO 9000:2015, which provides terms and definitions.
Noncompliance has legal consequences. Under § 820.10(e), failure to comply with an applicable requirement renders a device adulterated under section 501(h) of the FD&C Act.
Who Must Comply With 21 CFR 820?
The QMSR applies to manufacturers of finished devices intended for human use that are made in the United States or imported or offered for import into it. Companies performing only some operations must meet the requirements that apply to those operations.
Under § 820.3, a manufacturer includes any person who designs, manufactures, fabricates, assembles or processes a finished device. This includes roles such as:
- Contract manufacturers and contract sterilizers
- Specification developers
- Relabelers and repackers
- Remanufacturers
- Foreign manufacturers exporting to the US market
How does device class affect QMSR requirements?
- Class II and Class III: Design and development requirements (ISO 13485 Clause 7.3) apply.
- Class I, software-automated: Clause 7.3 also applies.
- Five listed Class I devices: Clause 7.3 applies to tracheobronchial suction catheters, non-powdered surgeon’s gloves, protective restraints, manual radionuclide applicator systems and radionuclide teletherapy sources.
- Exempt Class I devices: Some are exempt from CGMP in 21 CFR 862 to 892, but generally must still keep complaint files and meet § 820.35 records requirements.
Not covered: Manufacturers of components or parts of finished devices are outside Part 820, although FDA encourages them to consider its provisions. Not sure of your status? Start with FDA medical device registration and listing or ask your US FDA Agent.
How Is the QMSR Different From the QSR?
The main difference is structure. The QSR contained FDA’s own detailed quality system text. The QMSR uses ISO 13485:2016 as its base and adds a small number of FDA-specific provisions. Companies must also follow the changed terms, records and inspection approach.
| Area | Former QSR (to February 1, 2026) | QMSR (from February 2, 2026) |
|---|---|---|
| Core requirements | FDA text in Subparts A to O | ISO 13485:2016 by reference, plus Part 820 provisions |
| Definitions | FDA definitions in § 820.3 | ISO 13485 and ISO 9000 Clause 3, with listed exceptions; FD&C Act definitions take precedence |
| Key records | DHF, DMR, DHR and QSR | Design and development file, medical device file, device or batch record |
| Risk management | Explicit mainly in design validation | Explicit emphasis through ISO 13485 clauses across product realization |
| Additional FDA provisions | Within Subparts | § 820.10, § 820.35 (records) and § 820.45 (labeling and packaging) |
| Inspection method | QSIT | Compliance Program 7382.850 |
| Management review and audit records | Generally not reviewed by FDA | Available for FDA review |
The QMSR also keeps links to other FDA rules: UDI (Part 830), device tracking (Part 821), Medical Device Reporting (Part 803) and corrections and removals (Part 806).
Where Did the Old 21 CFR 820 Sections Go?
Former QSR section numbers are no longer current. Part 820 now contains § 820.1, § 820.3, § 820.7, § 820.10, § 820.35 and § 820.45. Other sections and Subparts C to O are reserved.
The table below helps teams who are used to the old numbers. It is a practical orientation by I3CGLOBAL, not an FDA crosswalk. FDA has no formal side-by-side comparison and expects manufacturers to perform their own gap analysis.
| Topic | Former QSR | Main reference now |
|---|---|---|
| Management responsibility | § 820.20 | ISO 13485 Clause 5 |
| Quality audit | § 820.22 | Clause 8.2.4 |
| Personnel | § 820.25 | Clause 6.2 |
| Design controls | § 820.30 | Clause 7.3; § 820.10(c) |
| Document controls | § 820.40 | Clause 4.2.4 |
| Purchasing controls | § 820.50 | Clause 7.4 |
| Identification and traceability | § 820.60, § 820.65 | Clauses 7.5.8, 7.5.9; Parts 821 and 830 |
| Production and process controls | § 820.70 | Clauses 6.3, 6.4, 7.5.1 |
| Inspection, measuring and test equipment | § 820.72 | Clause 7.6 |
| Process validation | § 820.75 | Clause 7.5.6 |
| Acceptance activities | § 820.80, § 820.86 | Clauses 7.4.3, 7.5.8, 8.2.6 |
| Nonconforming product | § 820.90 | Clause 8.3 |
| CAPA | § 820.100 | Clauses 8.5.2, 8.5.3 |
| Labeling and packaging | § 820.120, § 820.130 | § 820.45; Clauses 7.5.1, 7.5.11 |
| Handling, storage, distribution, installation | § 820.140 to § 820.170 | Clauses 7.5.1, 7.5.3, 7.5.11 |
| Records | § 820.180 to § 820.186 | Clauses 4.2.3, 4.2.5, 7.3.10, 7.5.1; § 820.35 |
| Complaint files | § 820.198 | Clauses 8.2.2, 8.2.3; § 820.35(a); Part 803 |
| Servicing | § 820.200 | Clause 7.5.4; § 820.35(b) |
| Statistical techniques | § 820.250 | Clauses 8.1, 8.4 |
Is ISO 13485 Certification Enough for the QMSR?
No. ISO 13485 certification helps, but it does not by itself ensure QMSR compliance. A certified company must still meet Part 820 and other applicable FDA requirements, and FDA will not accept a certificate in place of its own assessment.
Certified companies often have less to change, but they should check these areas:
- Complaint record content and FDA reportability decisions (§ 820.35 and Part 803)
- Servicing records (§ 820.35)
- UDI records and labeling checks (§ 820.35, § 820.45 and Part 830)
- Labeling and packaging controls to prevent mix-ups (§ 820.45)
- Corrections and removals (Part 806) and device tracking (Part 821)
- Whether FD&C Act definitions change how a procedure must be read
We support both routes together through ISO 13485 certification and MDSAP consulting. FDA accepts certain MDSAP audits as a substitute for its routine surveillance inspections.
Which FDA Requirements Apply on Top of ISO 13485?
Part 820 adds FDA-specific requirements for records, labeling, packaging, design scope and links to other FDA regulations. Meeting ISO 13485 alone does not address all of them, so check these items separately during a QMSR gap analysis.
What must complaint records include under § 820.35(a)?
Records of complaint review and investigation must document the evaluation of possible failures. If a similar complaint was already investigated, the manufacturer must record why another investigation is not needed. Required details include:
- Device name and date the complaint was received
- UDI or UPC, where applicable
- Complainant contact details
- Nature of the complaint and any corrective action
- Any reply to the complainant
Complaints that meet Part 803 criteria must also be reported to FDA. This is why complaint handling and Medical Device Reporting procedures should work together, with a clear decision record for each complaint.
What must servicing records include under § 820.35(b)?
Where servicing is a requirement, records must identify the device name, UDI or UPC, date of service, the person who performed it, the work carried out, and test and inspection data. Teams should check that service reports and field service software capture each item.
What does § 820.45 require for labeling and packaging?
Section 820.45 adds to ISO 13485 Clause 7.5.1. Manufacturers need documented procedures to keep labeling and packaging intact and properly inspected, stored and handled through processing, distribution and use. In practice, this means:
- Check labeling for accuracy before release or storage, including UDI or UPC, expiry date, and storage and handling instructions.
- Document the release of labeling for use.
- Organize labeling and packaging operations to prevent mix-ups.
- Inspect before use against the medical device file, and record the results.
Which other FDA regulations connect to the QMSR?
| FDA regulation | Subject | What to check in your QMS |
|---|---|---|
| 21 CFR Part 803 | Medical Device Reporting | Complaints are assessed for reportability and decisions are recorded |
| 21 CFR Part 806 | Corrections and removals | Advisory notice and recall procedures address FDA reporting |
| 21 CFR Part 821 | Device tracking | Tracking procedures exist where the device is subject to tracking |
| 21 CFR Part 830 | Unique Device Identification | UDI is assigned and recorded for each device or batch |
Which QSR Terms and Records Changed Under the QMSR?
Many procedures still use QSR language. The QMSR final rule does not require you to rename every document, but your system must clearly show how it meets current requirements. This table helps teams update their references.
| Former QSR term | Current QMSR reference | Practical action |
|---|---|---|
| Design History File (DHF) | Design and development file, Clause 7.3.10 | Confirm the file contains or references all design records |
| Device Master Record (DMR) | Medical device file, Clause 4.2.3 | Confirm current specifications and procedures are contained or referenced |
| Device History Record (DHR) | Medical device or batch record, Clause 7.5.1 | Confirm records support traceability and manufactured and released quantities |
| Quality System Record (QSR) | QMS documentation and records, Clause 4.2 | Update the document structure and cross-references |
A simple cross-reference between your old terms and the current requirements can help staff and investigators find information quickly. Keep it controlled like any other QMS document.
What Should You Check in a QMSR Gap Analysis?
A useful gap analysis looks beyond document titles and asks whether your records prove that the process works. Start with these questions:
- Regulatory references: Do procedures still cite repealed sections such as § 820.30 or § 820.100?
- Risk management: Are risk management processes documented across product realization, and do they influence decisions on design, suppliers, production and complaints?
- Management review: Do inputs and outputs meet ISO 13485 Clause 5.6, and could the records be shown to an FDA investigator?
- Internal audits: Is the audit program based on current requirements and process risk?
- Suppliers: Do your criteria go beyond a supplier’s certificate? FDA notes that certification may not give adequate assurance of supplier quality without further evaluation.
- Software: Are computer software used in the QMS and in production validated under the applicable ISO 13485 clauses?
- Feedback and complaints: Do these processes connect to CAPA, risk files and FDA reporting decisions?
- Training: Can staff explain the current process, not only the old one?
Manufacturers with an existing ISO 13485 certificate should not assume they have no gaps. Those with a QSR-only system usually need more mapping work, since the QMS may be organized around former FDA section numbers.
How Does FDA Inspect Under the QMSR?
FDA stopped using the Quality System Inspection Technique (QSIT) on February 2, 2026. It now inspects device manufacturers under Compliance Program 7382.850, Inspection of Medical Device Manufacturers.
Industry analyses of the program describe a risk-based approach. It is organized around six QMS areas: Management Oversight; Measurement, Analysis and Improvement; Design and Development; Change Control; Outsourcing and Purchasing; and Production and Service Provision. It also covers Medical Device Reporting, corrections and removals, tracking and UDI.
Prepare for these practical points:
- Risk files: Your risk management documents should be current and linked to your product and process decisions.
- Records now in scope: Management review, internal audit and supplier audit records should be ready for FDA review.
- No certificate: An FDA inspection does not end with a QMSR or ISO 13485 certificate.
- Observations: Inspection observations may be recorded on Form 483. Serious noncompliance may lead to a Warning Letter.
How Our FDA QMSR Consultants Can Help
I3CGLOBAL supports manufacturers at each stage of QMSR compliance. Scope depends on your device, sites and existing quality system.
QMSR gap analysis
We compare your QMS against Part 820 and ISO 13485:2016, then give you prioritized findings and a remediation plan.
QMS implementation and documentation
We draft or update your quality manual, procedures, work instructions, record forms, medical device file structure and CAPA process, using one system where it also needs to serve other markets.
QSR to QMSR remediation
If your system still reflects old QSR terms or section references, we help you update documents, records and traceability to current requirements.
Internal audits and training
We audit against current requirements and provide role-based training for staff, internal auditors and management.
Supplier qualification and audits
We help you set up supplier evaluation and controls under ISO 13485 Clause 7.4 and support supplier audits.
FDA inspection readiness
We carry out mock inspections and prepare your team for inspection against the current FDA approach.
Form 483 and Warning Letter response
We help prepare clear responses with root cause analysis, corrections, corrective action plans and evidence of follow-up.
One QMS for several markets
We can design one documentation structure for FDA, ISO 13485 and other market requirements, so your team avoids maintaining separate systems.
Ongoing QMS support
We can provide continued support for internal audits, training and regulatory updates after your initial QMSR project ends.
Need a cost estimate? See our QMSR consulting fee page. If you are also preparing a submission, see FDA 510(k) support.
Step by step
How Do You Move to QMSR Compliance?
These seven steps help you move an existing quality system to current FDA requirements. Each step should leave evidence your team can show during an inspection.
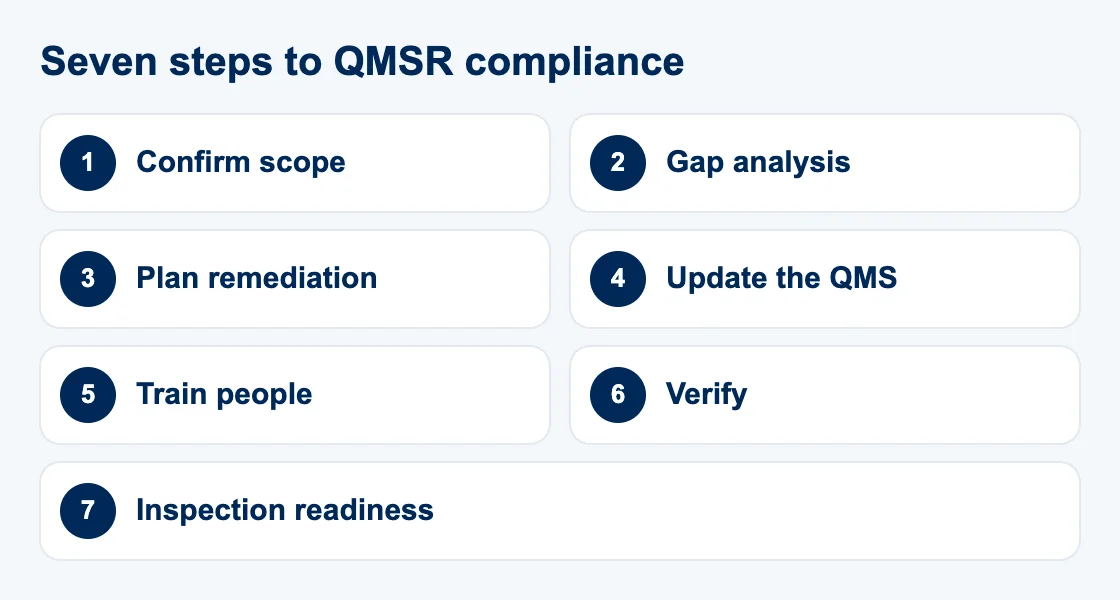
- Confirm scopeIdentify your devices, classes, CGMP exemptions, activities and sites.
- Run a gap analysisCompare your QMS with ISO 13485:2016 and Part 820, including FDA-specific provisions.
- Plan remediationAssign owners, priorities and evidence requirements, starting with the highest product risk.
- Update the QMSRevise procedures, terminology, records, risk management links and supplier controls.
- Train peopleMake sure each role understands what changed in its work.
- VerifyRun internal audits and management review, then address findings through CAPA.
- Prepare for inspectionHold a mock inspection and keep records ready for FDA review.
Why Choose I3CGLOBAL’s 21 CFR 820 Consultants?
Combined US and international scope
We work across FDA, ISO 13485, EU and UK requirements, so one quality system can support several markets.
Offices in several regions
I3CGLOBAL lists offices in the United States, United Kingdom, Germany, Portugal, India, South Korea and Malaysia.
Medical device focus
Regulatory consulting for medical devices, IVDs and life sciences is our core work.
Practical deliverables
We aim for documents your team can use daily and explain during an inspection.
Ongoing support
We can continue with audits, training and regulatory updates after your project ends.
Learn more about I3CGLOBAL and our team. No consultant can guarantee an FDA inspection outcome; compliance remains the manufacturer’s responsibility.
Work with I3CGLOBAL
Get Your QMSR Gaps Reviewed by FDA QMSR Consultants
Our regulatory team can review your quality system against 21 CFR Part 820 and ISO 13485:2016, then help you close the gaps.
- QMSR gap analysis and remediation plan
- Procedures, records and medical device file updates
- Internal audits and mock FDA inspections
- Form 483 and Warning Letter response support
Request your QMSR proposal
Share your device and current QMS status. You can also review our QMSR consulting fees.
FAQs
FDA QMSR and 21 CFR 820: Frequently Asked Questions
What is 21 CFR 820 now?
21 CFR Part 820 is the FDA regulation that sets current good manufacturing practice requirements for finished medical devices. Since February 2, 2026, it has been titled the Quality Management System Regulation (QMSR) and incorporates ISO 13485:2016 by reference.
Is the QSR still in force?
No. The QMSR amended and replaced the QSR text in Part 820 on February 2, 2026. Most former sections, such as § 820.30 and § 820.100, are now reserved or removed. Their topics are covered mainly through ISO 13485:2016 clauses.
Is the QMSR the same as ISO 13485?
No. The QMSR incorporates ISO 13485:2016 and adds FDA-specific provisions in Part 820, including requirements for complaint and servicing records, UDI and labeling controls. Where ISO 13485 conflicts with the FD&C Act or its regulations, the Act and regulations control.
Does ISO 13485 certification mean a company complies with the QMSR?
Not by itself. FDA states that certification alone does not ensure QMSR compliance, and that it will not accept ISO 13485 certificates in place of its own assessment. A certified company should still check FDA-specific requirements.
Does ISO 13485 certification replace an FDA inspection?
No. FDA keeps its inspection authority and carries out its own assessment. Certain MDSAP audits may substitute for FDA routine surveillance inspections, but that does not exempt a company from all FDA inspections.
Does the FDA issue a QMSR certificate?
No. FDA inspections do not result in a QMSR certificate or an ISO 13485 certificate of conformity. ISO 13485 certificates are issued by certification bodies.
Are Class I devices subject to the QMSR?
Many are, but some Class I devices are exempt from CGMP requirements by their classification regulations in 21 CFR 862 to 892. Exempt manufacturers must generally still keep complaint files and meet the records requirements of § 820.35. Check the exact classification regulation for your device.
Do Class I devices need design controls?
Most do not. Design and development requirements apply to Class II and Class III devices, Class I devices automated with computer software, and five Class I device types listed in § 820.10(c).
Do foreign manufacturers have to follow the QMSR?
Yes, when their finished devices are imported or offered for import into the United States. Devices that appear adulterated may be refused admission under section 801(a) of the FD&C Act.
Do component manufacturers have to comply?
Part 820 does not apply to manufacturers of components or parts of finished devices, although FDA encourages them to consider its provisions as appropriate. Finished device manufacturers remain responsible for controlling their suppliers under ISO 13485 purchasing requirements.
What happened to the DHF, DMR and DHR?
FDA did not keep separate requirements for these record types. Their content is now expected mainly in the design and development file (Clause 7.3.10), medical device file (Clause 4.2.3) and medical device or batch record (Clause 7.5.1).
Is ISO 14971 required under the QMSR?
FDA did not incorporate ISO 14971 into the QMSR. However, ISO 13485 requires documented risk management processes in product realization, and ISO 14971 is the recognized risk management standard many manufacturers use to meet this requirement.
What do FDA investigators review during a QMSR inspection?
FDA stopped using QSIT on February 2, 2026 and now uses Compliance Program 7382.850, Inspection of Medical Device Manufacturers. Investigators may also review management review, internal audit and supplier audit records, which were generally not reviewed under the former QSR.
What is an FDA Form 483?
Form 483 lists inspection observations that an FDA investigator believes may be violations. It is issued at the end of an inspection. Companies should respond in writing with root cause analysis, corrections and a realistic corrective action plan.
How long does a QMSR transition take?
There is no fixed period. It depends on your device class, your current QMS, ISO 13485 status, number of sites and how much documentation must change. A gap analysis gives the realistic basis for your plan.