
Class Is Medical Device (Sterile Class I) under EU MDR
A Class Is medical device, also written Class 1s, is a Class I device placed on the market in sterile condition. It is still low risk, but under Article 52(7) of EU MDR 2017/745 a Notified Body must assess how the manufacturer establishes, secures and maintains sterility. All other aspects stay under the manufacturer’s own Class I declaration.
Classification rules
Which Rules Make a Device Class Is?
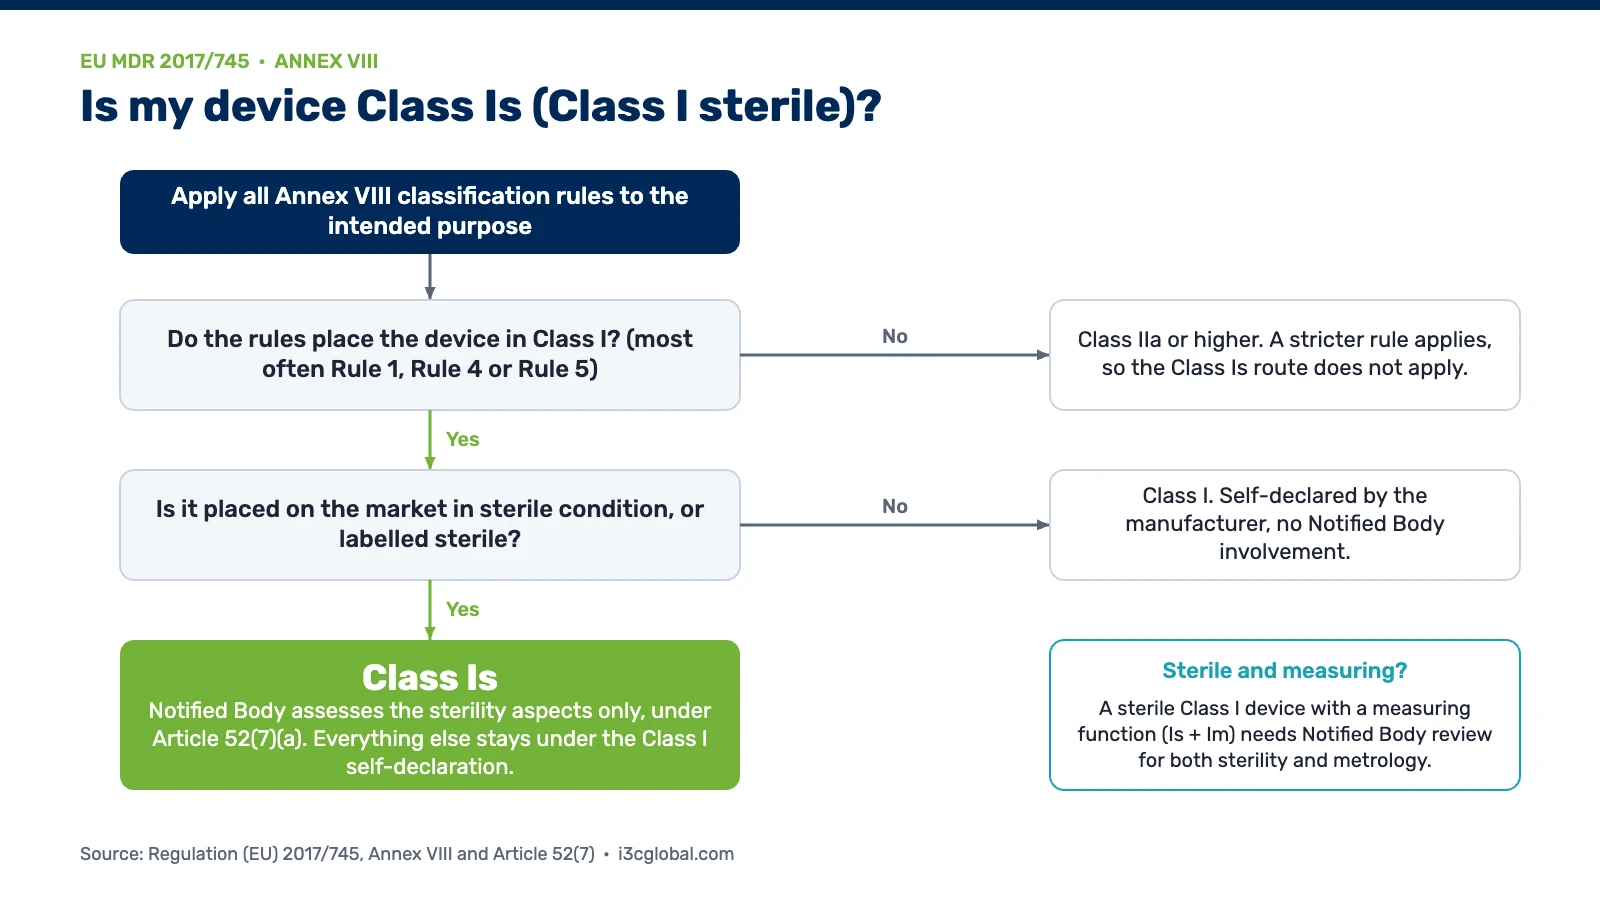
No Annex VIII rule is called “Class Is” (the Commission classification guidance is MDCG 2021-24). A device is Class Is when two things are true:
- The classification rules place it in Class I, most often under Rule 1, Rule 4 or Rule 5.
- The manufacturer places it on the market sterile, or claims it is sterile on the label.
| Rule | Class I condition | Becomes Class Is when |
|---|---|---|
| Rule 1 | Non-invasive device | Supplied sterile |
| Rule 4 | Injured-skin device acting as a mechanical barrier, for compression or absorbing exudates | Supplied sterile |
| Rule 5 | Transient use through a body orifice, or short-term use in the oral cavity as far as the pharynx, in the ear canal up to the ear drum, or in the nasal cavity | Supplied sterile |
If a stricter rule applies, for example because the device is surgically invasive, the device is Class IIa or higher and the Class Is route does not apply.
Examples
What Are Examples of Class Is Medical Devices?
These examples are typical. The final class always depends on the intended purpose and claims.
| Device (typical) | Rule | Why it is usually Class Is |
|---|---|---|
| Sterile gauze swabs and pads for absorbing exudate | Rule 4 | Mechanical barrier and absorption, supplied sterile. |
| Sterile non-medicated dressings for minor wounds | Rule 4 | No claim to manage the wound micro-environment. |
| Sterile examination gloves | Rule 1 / Rule 5 | Intact skin or transient orifice use, supplied sterile. |
| Sterile urine collection bags | Rule 1 | Non-invasive collection, supplied sterile. |
| Sterile single-use specula for transient use | Rule 5 | Transient use through a body orifice, supplied sterile. |
CE marking route
How Do You CE Mark a Class Is Device?
Article 52(7)(a) lets the manufacturer choose either Annex IX Chapters I and III (quality management system assessment) or Annex XI Part A (production quality assurance). In both cases the Notified Body’s review is limited to sterility.
- Confirm Class I and the sterile claimApply all Annex VIII rules and record why the device is Class I.
- Choose and validate the sterilisation methodValidate the process with installation, operational and performance qualification (IQ, OQ, PQ). See our sterilisation validation service.
- Validate the sterile barrier systemValidate packaging, sealing and shelf life to EN ISO 11607-1 and 11607-2 (packaging validation).
- Build the QMS around sterilityCover cleanroom or controlled environment, bioburden, routine release and supplier control for contract sterilisers.
- Complete the technical documentationPrepare the full Annex II and III technical documentation, clinical evaluation report and GSPR checklist, as for any Class I device.
- Apply to a Notified BodyChoose a Notified Body whose designation covers sterile devices (check the EU Notified Body database) and the right conformity assessment annex.
- Pass the Notified Body assessmentThe Notified Body reviews sterility documentation and audits the related QMS processes.
- Sign the declaration and affix the CE markAdd the Notified Body number next to the CE mark. Register as an actor in EUDAMED to obtain your SRN before placing the device on the market, then register the device (Basic UDI-DI and UDI-DI). Use of the first four EUDAMED modules has been mandatory since 28 May 2026. See our EUDAMED registration service.
Sterility evidence
What Sterilisation and Packaging Evidence Does Class Is Need?
Choose the sterilisation method that suits the device materials and design. The table lists the main harmonised standards.
| Method | Standard | Typical use |
|---|---|---|
| Ethylene oxide (EO) | EN ISO 11135 | Heat-sensitive devices and complex shapes. Needs aeration and EO residue testing to EN ISO 10993-7. |
| Radiation (gamma, electron beam, X-ray) | EN ISO 11137 | Single-use devices in final packs. Check material stability at the chosen dose. |
| Moist heat (steam) | EN ISO 17665-1 | Heat-stable materials such as metals and some textiles. |
| Sterile barrier packaging | EN ISO 11607-1 and 11607-2 | Packaging design, sealing validation and shelf-life testing for every method. |
Validation reports
IQ, OQ and PQ protocols and reports with a clear conclusion and raw data.
Routine control
Batch release records, biological or dosimetric indicators and bioburden monitoring.
Sterile labelling
The sterile symbol with the method, expiry date and “do not use if package is damaged” information.
Clinical evaluation
A clinical evaluation report under Article 61, as for every Class I device.
After CE marking. Your obligations continue. Keep the post-market surveillance plan running, update the PMS report under Article 85 when necessary, and report serious incidents and field safety corrective actions through the vigilance system. The Notified Body carries out surveillance audits of the sterilisation and packaging processes during the certificate’s validity, and it must be told about significant changes to those processes before you implement them. Manufacturers outside the EU also need an EU authorised representative (Article 11), and every manufacturer needs a person responsible for regulatory compliance (Article 15). Keep the technical documentation available for at least 10 years after the last device is placed on the market, or 15 years for implantable devices. The Notified Body certificate is valid for a maximum of five years and is renewed after reassessment (Article 56(2)).
Borderline cases
What Are Common Class Is Mistakes?
- Assuming every sterile device is Class Is. Sterile surgically invasive devices, such as single-use scalpels or sterile surgical gloves, are usually Class IIa under Rule 6.
- Weak packaging evidence. A validated sterilisation cycle is not enough without seal strength, integrity and shelf-life data.
- Unqualified contract sterilisers. The manufacturer stays responsible. The steriliser must be controlled as a critical supplier.
- Wound claims. A sterile dressing that claims to promote healing or manage moisture is usually Class IIa, not Class Is.
- Missing EO residue data. EO-sterilised devices need residue limits shown under EN ISO 10993-7.
Compare
How Does Class Is Compare with Other Classes?
| Class | Notified Body scope |
|---|---|
| Class I | None, self-declared |
| Class Is | Sterile conditions only |
| Class Im | Metrological requirements only |
| Class Ir | Reuse aspects only |
| Class IIa and above | Full QMS and technical documentation sampling |
See all classes side by side on Class I vs IIa vs IIb vs III.
Work with I3CGLOBAL
Certify Your Sterile Class I Device
I3CGLOBAL has completed 300+ CE certifications, with offices in India, Europe and the USA. We prepare Class Is files that Notified Bodies can review quickly.
- Sterilisation and packaging validation review
- Technical documentation and GSPR checklist
- Clinical evaluation report
- Notified Body coordination
Request your proposal
Send us your device description and sterilisation method, and we will prepare a Class Is proposal.
FAQs
Class Is Medical Device: Frequently Asked Questions
Is Class 1s the same as Class Is?
Yes. Class Is and Class 1s are two ways of writing the same category: Class I medical devices placed on the market in sterile condition. The MDR text uses Class I with the word sterile, and industry shortens it to Is.
Does a Class Is device need a Notified Body?
Yes. Under Article 52(7)(a) of the MDR, a Notified Body must assess the aspects of manufacture that establish, secure and maintain sterile conditions. The rest of the device remains under the manufacturer’s Class I self-declaration.
What does the Notified Body review for Class Is?
It reviews sterilisation validation, routine process control, sterile barrier packaging validation, the related parts of the quality management system and the sterile labelling. It does not issue a full technical documentation review of non-sterile aspects.
Which sterilisation standards apply to Class Is devices?
The most used harmonised standards are EN ISO 11135 for ethylene oxide, EN ISO 11137 for radiation and EN ISO 17665 for moist heat. EN ISO 11607 covers sterile barrier packaging. Choose the method that suits the device materials.
Is a sterile device always Class Is?
No. A sterile device is Class Is only if the classification rules give Class I. Many sterile devices, such as surgically invasive instruments or wound dressings for deep wounds, are Class IIa or higher because a stricter rule applies.
Can a Class Is device also be Class Im?
Yes. A sterile Class I device with a measuring function needs Notified Body involvement for both sterility and metrology. The certificate scope then covers both aspects.
What certificate does a Class Is manufacturer receive?
The Notified Body issues an EU quality management system certificate under Annex IX (Chapters I and III) or an EU quality assurance certificate under Annex XI Part A. Its scope is limited to the aspects of manufacture that establish, secure and maintain sterile conditions, and it is valid for up to five years (Article 56(2)). The manufacturer then signs the EU declaration of conformity for the whole device.
How long does Class Is CE marking take?
There is no fixed legal timeline. The duration depends on how complete your sterilisation and packaging validation is, the Notified Body’s capacity and the conformity assessment annex you choose. Having IQ, OQ and PQ reports, EN ISO 11607 packaging data and sterile labelling ready before you apply is the biggest factor you control. Contact I3CGLOBAL for a device-specific estimate.
Is a Class Is certificate recognised in the UK or the USA?
No. The EU certificate is not transferable. In Great Britain the device must meet the UK Medical Devices Regulations 2002, where a UK Approved Body assesses the sterility aspects for UKCA marking; CE marked devices are accepted in Great Britain only during the transitional period set in UK law. In the USA the FDA classifies devices by product code, not by the EU class, so a Class Is device may be exempt, need a 510(k) or follow another pathway. See our UKCA marking and FDA 510(k) pages.