Class III Medical Device (Class 3) under EU MDR
A Class III medical device is in the highest risk class under EU MDR 2017/745. Annex VIII places a device in Class III when it is in direct contact with the heart, central circulatory system or central nervous system, is an active implant, contains an ancillary medicinal substance, or meets another high-risk rule. Every Class III device needs a Notified Body for CE marking, and most need a clinical investigation under Article 61(4).
Key facts: Risk class: highest of four (I, IIa, IIb, III). Classification: Annex VIII, where 12 of the 22 rules can lead to Class III. Conformity assessment: Annex IX, or Annex X with Annex XI (Article 52(3)). Clinical evidence: clinical investigation normally required; SSCP and annual PSUR mandatory.
Classification rules
Which Rules Make a Device Class III?
Class III comes from applying the 22 rules in Annex VIII of the MDR to the device’s intended purpose. If more than one rule applies, the strictest rule decides the class. The table lists the rules that lead to Class III. See all 22 rules on the medical device classification page. The European Commission guidance MDCG 2021-24 explains how to apply the rules.
Annex VIII rules that lead to Class III
| Rule | When the device is Class III |
|---|---|
| Rule 3 | Non-invasive devices made of a substance used in vitro in direct contact with human cells, tissues, organs or embryos before they are implanted or administered |
| Rule 6 | Transient surgically invasive devices that control, diagnose, monitor or correct a defect of the heart or central circulatory system through direct contact, or that are in direct contact with the heart, central circulatory system or central nervous system |
| Rule 7 | Short-term surgically invasive devices in direct contact with the heart, central circulatory system or central nervous system, or that have a biological effect or are wholly or mainly absorbed |
| Rule 8 | Implantable and long-term surgically invasive devices in direct contact with the heart, central circulatory system or central nervous system; active implantables and their accessories; breast implants; surgical meshes; total and partial joint replacements; spinal disc replacements and implants in contact with the spinal column; devices with a biological effect, wholly or mainly absorbed, that undergo chemical change in the body (except in the teeth) or that administer medicines |
| Rule 9 | Active devices that control, monitor or directly influence the performance of active implantable devices |
| Rule 11 | Software providing information used for decisions that may cause death or an irreversible deterioration of health |
| Rule 14 | Devices that include, as an integral part, a medicinal substance with an ancillary action, including a human blood or plasma derivative |
| Rule 15 | Contraception or STD-prevention devices that are implantable or long-term invasive |
| Rule 18 | Devices made with non-viable human or animal tissues or cells, or their derivatives (except animal-tissue devices for contact with intact skin only) |
| Rule 19 | Devices containing nanomaterial with high or medium potential for internal exposure |
| Rule 21 | Substances introduced into the body that are systemically absorbed to achieve their purpose, or that act in the stomach or lower gastrointestinal tract and are systemically absorbed |
| Rule 22 | Active therapeutic devices with an integrated diagnostic function that significantly determines patient management, such as closed-loop systems and automated external defibrillators |
Software and accessories
Software that drives or influences a device falls into the same class as the device. Accessories are classified in their own right (Annex VIII, implementing rules 3.3 and 3.2).
Examples
What Are Examples of Class III Medical Devices?
The examples below are typical. The class of your device always depends on its intended purpose and claims, so confirm the rule for your own product.
| Device | Rule | Why it is Class III |
|---|---|---|
| Prosthetic heart valve | Rule 8 | Implant in direct contact with the heart |
| Coronary stent | Rule 8 | Implant in direct contact with the central circulatory system |
| Drug-eluting stent | Rules 8 and 14 | Implant in the circulatory system with an ancillary medicinal substance |
| Implantable pacemaker or neurostimulator | Rule 8 | Active implantable device |
| Pacemaker programmer | Rule 9 | Active device that controls an active implantable device |
| Hip or knee joint replacement | Rule 8 | Total or partial joint replacement |
| Spinal disc replacement | Rule 8 | Implant in contact with the spinal column |
| Breast implant | Rule 8 | Named in Rule 8 |
| Surgical mesh | Rule 8 | Named in Rule 8 |
| Absorbable suture | Rule 8 | Long-term implant that is wholly or mainly absorbed |
| Collagen or bovine-tissue implant | Rule 18 | Made from non-viable animal tissue |
| IVF culture media | Rule 3 | In direct contact with human embryos before implantation |
| Automated external defibrillator | Rule 22 | Integrated diagnosis that decides the therapy |
Note: the ancillary screws, wedges, plates and instruments used with joint and spinal implants are not automatically Class III. Classify each component on its own.
CE marking
How Do You CE Mark a Class III Device?
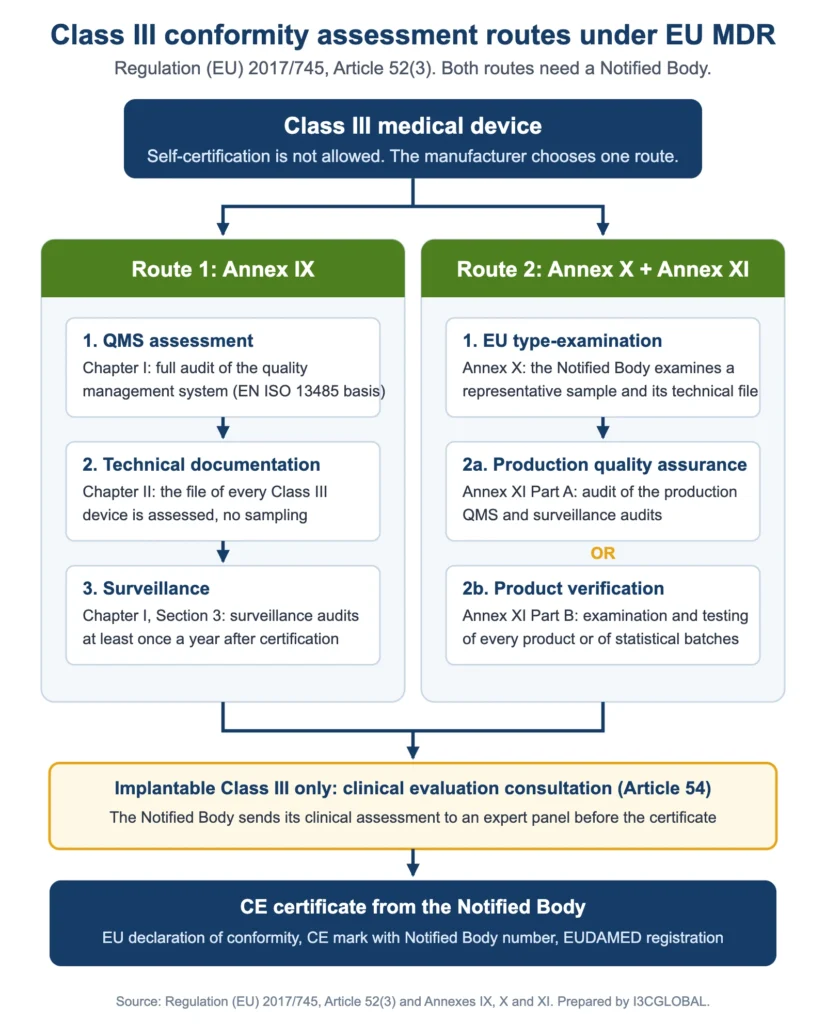
A Class III device cannot be self-certified. Article 52(3) gives two conformity assessment routes, and both involve a Notified Body.
Route 1: Annex IX
Full quality management system assessment (Chapter I), assessment of the technical documentation of every device (Chapter II), and ongoing surveillance audits (Chapter I, Section 3).
Route 2: Annex X with Annex XI
EU type-examination of a representative sample (Annex X), followed by production quality assurance or product verification (Annex XI).
- Confirm the class and ruleRecord the Annex VIII rule and your reasoning in the technical documentation.
- Set up the QMSImplement an MDR quality management system, usually to EN ISO 13485, with risk management to EN ISO 14971. Software devices also follow IEC 62304, and clinical investigations follow ISO 14155.
- Build the clinical evidencePlan the clinical evaluation and the clinical investigation required by Article 61(4), unless a justified exemption applies.
- Prepare the technical documentationCompile the technical documentation: Annex II and Annex III files, including GSPR checklist, verification and validation, labelling and the PMS plan.
- Notified Body reviewThe Notified Body (listed in the European Commission NANDO database) audits the QMS and reviews the technical documentation. For implantable Class III devices it also runs the clinical evaluation consultation procedure.
- CE certificate, declaration and registrationSign the EU declaration of conformity, affix the CE mark with the Notified Body number, and register the device and UDI in EUDAMED (mandatory since 28 May 2026).
Clinical evaluation consultation procedure (CECP): under Article 54, the Notified Body sends its clinical evaluation assessment report for Class III implantable devices to an expert panel. The panel may give a scientific opinion, and the Notified Body must give due consideration to it before issuing the certificate.
The consultation is not required for certificate renewals, for modifications of a device the same manufacturer already markets for the same intended purpose, or where common specifications for the clinical evaluation of that device type exist (Article 54(2)).
Documentation
What Technical Documentation and Clinical Evidence Does Class III Need?
All devices need technical documentation and a clinical evaluation. For Class III, the Notified Body checks the evidence for each device, and several extra documents apply.
Clinical investigation
Article 61(4) requires clinical investigations for implantable and Class III devices. Exemptions are narrow, for example some device modifications by the same manufacturer and some equivalent devices with full access to data.
Summary of safety and clinical performance
Article 32 requires an SSCP for implantable and Class III devices. The Notified Body validates it and it is published in EUDAMED.
PSUR every year
Article 86 requires a periodic safety update report, updated at least annually for Class IIb and Class III devices. Class III manufacturers submit it to their Notified Body, which reviews it and records its evaluation. Submission will move to EUDAMED once the vigilance and post-market surveillance module becomes mandatory; as of November 2026 it is not.
Implant card
Article 18 requires an implant card and patient information for implantable Class III devices. Non-implantable Class III devices do not need one. Sutures, staples, dental fillings, dental braces, tooth crowns, screws, wedges, plates, wires, pins, clips and connectors are exempt.
PMCF
A post-market clinical follow-up plan and report keep the clinical evidence current throughout the device’s life.
UDI
Class III and implantable devices have carried the UDI on their labels since 26 May 2021.
Other obligations and transition rules
Every manufacturer also needs a person responsible for regulatory compliance (PRRC). Manufacturers outside the EU must appoint an EU authorised representative (Article 11).
Legacy MDD devices: under Regulation (EU) 2023/607, Class III devices with an MDD certificate that was valid on 26 May 2021 may stay on the market until 31 December 2027 if the device still complies with the MDD, has no significant change in design or intended purpose, presents no unacceptable risk to health or safety, the manufacturer had an MDR-compliant QMS in place by 26 May 2024, and a formal application was lodged with a Notified Body by 26 May 2024 with a written agreement signed by 26 September 2024. See the MDR transition timeline.
Regulatory status (November 2026): on 16 December 2025 the European Commission proposed a targeted revision of the MDR and IVDR. It is still a proposal before the European Parliament and Council. The requirements on this page reflect the MDR as currently in force; this page will be updated if the amendment is adopted.
Borderline cases
What Are Common Class III Classification Mistakes?
- Calling every implant Class III. Most implants are Class IIb under Rule 8. They become Class III only through a specific condition such as heart contact, absorption or a named device type.
- Missing an ancillary medicine. A coating with a medicinal substance that supports the device moves it to Class III under Rule 14, even when the base device is lower risk.
- Overlooking animal-derived materials. Collagen, gelatin and other non-viable animal tissue derivatives usually trigger Rule 18.
- Classifying software on its own when it drives a device. Software that controls a Class III device takes the device’s class.
- Assuming MDD classes still apply. The MDR moved several device types up to Class III, including joint replacements, spinal implants, breast implants and surgical meshes.
- Planning equivalence without data access. For Class III, claiming equivalence to another manufacturer’s device usually requires a contract giving full access to its technical documentation.
Class IIb vs Class III
What Is the Difference Between Class IIb and Class III?
Both classes need a Notified Body, but Class III adds a review of every device file, a clinical investigation for all devices and an SSCP. For the full side-by-side view of all four classes, see Class I vs IIa vs IIb vs III.
| Requirement | Class IIb | Class III |
|---|---|---|
| Notified Body | Required | Required |
| Technical documentation review | At least one representative device per generic device group (Annex IX, Section 2.3) | Every device (Annex IX, Chapter II) |
| Clinical investigation | Required for implantable Class IIb devices (Article 61(4)); otherwise based on the clinical evaluation | Required for all Class III devices unless an Article 61(4) to 61(6) exemption applies |
| Clinical evaluation consultation (CECP) | Only active devices intended to administer or remove a medicinal product (Rule 12) | All implantable Class III devices (Article 54) |
| Summary of safety and clinical performance (SSCP) | Implantable Class IIb devices only | All Class III devices (Article 32) |
| PSUR | At least annually (Article 86) | At least annually, submitted to the Notified Body (Article 86) |
Lower-risk classes: Class I is low risk and self-declared (except Is, Im and Ir), and Class IIa is medium risk with a Notified Body required.
Work with I3CGLOBAL
Class III CE Marking Consulting from I3CGLOBAL
I3CGLOBAL has completed 300+ CE certifications worldwide, with offices in India, Europe and the USA. Tell us about your device and we will suggest the right route.
Our MDR CE marking case studies include Class III projects such as an insulin pump, a defibrillator and an intrauterine device classified under Rule 14.
- Classification and conformity route
- Technical documentation and GSPR checklist
- Clinical evaluation, CER and PMCF
- Notified Body submission and review responses
Request your proposal
Share your device details and we will prepare an MDR proposal.
FAQs
Class III Medical Device: Frequently Asked Questions
Is Class 3 the same as Class III?
Yes. The MDR writes the class in Roman numerals as Class III. Class 3 is the same class written with an Arabic number. Both refer to the highest-risk class under Annex VIII of EU MDR 2017/745.
Does every Class III device need a clinical investigation?
Usually yes. Article 61(4) requires clinical investigations for implantable and Class III devices. Limited exemptions exist, for example some modified devices from the same manufacturer, some equivalent devices with full data access, and certain well-established technologies listed in Article 61(6).
What is the clinical evaluation consultation procedure (CECP)?
CECP is the Article 54 procedure for Class III implantable devices. The Notified Body sends its clinical evaluation assessment report to an expert panel, which may give a scientific opinion. The Notified Body must consider that opinion before issuing the certificate.
How often must a Class III PSUR be updated?
At least once a year. Under Article 86, Class III manufacturers update the periodic safety update report annually and submit it to their Notified Body for review. When the EUDAMED vigilance module becomes mandatory, submission will take place through EUDAMED.
Can software be a Class III medical device?
Yes. Under Rule 11, software is Class III when the information it provides is used for decisions that may cause death or an irreversible deterioration of a person’s health. Software that drives a Class III device also takes the device’s class.
Can a Class III device be self-certified?
No. A Class III device always needs a Notified Body. The Notified Body assesses the quality management system and the technical documentation of every Class III device before the CE certificate is issued.
How long does Class III CE marking take?
There is no fixed timeline. The longest steps are usually the clinical investigation, compiling the technical documentation and the Notified Body review of the QMS and every device file. For implantable Class III devices, the expert panel must decide within 21 days whether to give an opinion and deliver it within 60 days (Annex IX, Section 5.1). Overall duration depends on the device, the clinical data available and Notified Body capacity.
Is EU Class III the same as US FDA Class III?
No. The EU and US systems classify devices with different rules. In the EU, Class III follows the Annex VIII rules of MDR 2017/745 and leads to Notified Body assessment. In the US, Class III under 21 CFR Part 860 generally requires a Premarket Approval (PMA). A device can be Class III in one system and a lower class in the other, so each market needs its own classification.
Does a Class III manufacturer outside the EU need an EU Authorised Representative?
Yes. Article 11 requires every manufacturer not established in an EU Member State to appoint a single EU Authorised Representative before placing a device on the EU market, whatever the class. The representative is named on the label and must have its own person responsible for regulatory compliance.
Do all Class III devices need an implant card?
No. Article 18 applies to implantable devices only. A Class III device that is not implanted, such as Class III software or an IVF culture medium, does not need an implant card. Implantable Class III devices need one unless they are on the exemption list (sutures, staples, screws, plates and similar components).